 filings.tools
Item 2. Management’s Discussion and Analysis of Financial Condition and Results of Operations
59K characters. Original on sec.gov · Markdown
Item 2. Management’s Discussion and Analysis of Financial Condition and Results of Operations
OVERVIEW
We are a global biotechnology company that invests in scientific innovation to create transformative medicines for people with serious diseases, with a focus on specialty markets. We have four approved medicines that treat the underlying cause of cystic fibrosis (“CF”), a life-threatening genetic disease, and one approved therapy that treats severe sickle cell disease (“SCD”) and transfusion dependent beta thalassemia (“TDT”), life-shortening inherited blood disorders. Our pipeline includes clinical-stage programs in CF, sickle cell disease, beta thalassemia, acute and neuropathic pain, APOL1-mediated kidney disease, type 1 diabetes, myotonic dystrophy type 1, alpha-1 antitrypsin deficiency, and autosomal dominant polycystic kidney disease.
Our four approved CF medicines, led by TRIKAFTA/KAFTRIO (elexacaftor/tezacaftor/ivacaftor and ivacaftor), are being used to treat nearly three quarters of the approximately 92,000 people with CF in North America, Europe, and Australia. We are evaluating our CF medicines in additional patient populations, including younger children, with the goal of having small molecule treatments for all people who have at least one mutation in their cystic fibrosis transmembrane conductance regulator (“CFTR”) gene that is responsive to our CFTR modulators. We also are pursuing messenger ribonucleic acid (“mRNA”) and genetic therapies for people with CF who do not make full-length CFTR protein and, as a result, cannot benefit from our current CF medicines.
In late 2023 and early 2024, CASGEVY (exagamglogene autotemcel or “exa-cel”), an ex-vivo, non-viral CRISPR/Cas9 gene-edited cell therapy, was approved in the U.S., the E.U., the United Kingdom (“U.K.”), the Kingdom of Saudi Arabia (“Saudi Arabia”), and the Kingdom of Bahrain (“Bahrain”) for the treatment of people 12 years of age and older with SCD or TDT. We estimate approximately 35,000 people with severe SCD or TDT could be eligible for CASGEVY in the U.S. and Europe, with additional people in Saudi Arabia and Bahrain. In addition, we are preparing for near-term launches of potential new products in CF and acute pain.
Financial Highlights
| Revenues | In the first quarter of 2024, our net CF product revenues increased to $2.7 billion as compared to $2.4 billion in the first quarter of 2023. The increase was primarily due to the continued strong uptake of TRIKAFTA/KAFTRIO in ex-U.S. markets with recently achieved reimbursements and label extensions in younger age groups and performance of TRIKAFTA in the U.S., following the launch of TRIKAFTA in children with CF 2 to 5 years of age. | ||||
| Expenses | Our total research and development (“R&D”), acquired in-process research and development (“AIPR&D”), and selling, general and administrative (“SG&A”) expenses decreased to $1.2 billion in the first quarter of 2024 as compared to $1.3 billion in the first quarter of 2023. The decrease was primarily due to decreased AIPR&D. Cost of sales was 13% in the first quarter of 2024 as compared to 11% in the first quarter of 2023, primarily due to cost of sales associated with CASGEVY. | ||||
| Cash | Our total cash, cash equivalents and marketable securities increased to $14.6 billion as of March 31, 2024 as compared to $13.7 billion as of December 31, 2023 primarily due to our cash from operations driven by our net product revenues, partially offset by business development payments and repurchases of our common stock. |
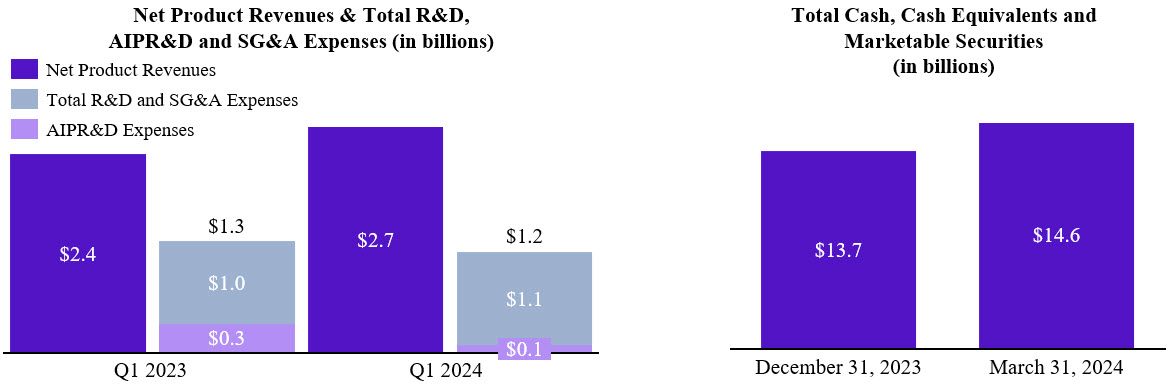
Note: Charts above may not add due to rounding.
Business Updates
Marketed Products
Cystic Fibrosis
We expect to grow our CF business with (i) continued uptake by patients in countries where we are early in our launch, (ii) label expansions, including into younger patient groups, and (iii) growth in the number of people living with CF. Recent progress in activities supporting continued uptake and label expansions is included below:
- The European Commission approved KALYDECO for the treatment of infants with CF from 1 month to less than 4 months of age with specific mutations in the CFTR gene.
Sickle Cell Disease and Beta Thalassemia
-
CASGEVY is approved in the U.S., the E.U., the U.K., Saudi Arabia, and Bahrain for people 12 years of age and older with SCD or TDT.
-
We completed regulatory submissions for CASGEVY in both SCD and TDT in Switzerland and Canada, and our regulatory submission in Canada has been granted priority review.
-
We have activated more than 25 authorized treatment centers globally, and multiple patients have initiated cell collection.
-
We entered into multiple agreements with commercial and government health insurance providers in the U.S. to provide access to CASGEVY. We have also secured reimbursed access for people with SCD or TDT in Saudi Arabia and Bahrain, as well as for people with TDT in France through an early access program.
Potential Near-Term Launch Opportunities
We are preparing for the following near-term launches of potential new products:
Vanzacaftor/tezacaftor/deutivacaftor in CF
-
In February 2024, we announced positive results from the pivotal program for our next-generation triple combination of vanzacaftor/tezacaftor/deutivacaftor (the “vanzacaftor triple”) for people with CF 6 years of age and older.
-
We submitted regulatory marketing applications for the vanzacaftor triple in people with CF 6 years of age and older to the U.S. Food and Drug Administration (“FDA”), using a priority review voucher, and to the European Medicines Agency (the “EMA”). We expect to complete additional global regulatory submissions for people with CF 6 years of age and older in 2024.
Suzetrigine in Acute Pain
-
In January 2024, we announced positive results from our Phase 3 clinical trials evaluating our lead compound, suzetrigine (formerly VX-548), a selective NaV1.8 inhibitor for the treatment of moderate-to-severe acute pain.
-
In the U.S., the FDA has granted rolling submission to the new drug application (“NDA”) for suzetrigine for the treatment of moderate-to-severe acute pain and we have begun the NDA submission process. We expect to complete the submission in the second quarter of 2024. Suzetrigine has been granted Fast Track and Breakthrough Therapy designations by the FDA in moderate-to-severe acute pain.
Pipeline
We continue to advance a diversified pipeline of potentially transformative medicines for serious diseases utilizing a range of modalities. Recent and anticipated progress in activities supporting these efforts is included below:
Cystic Fibrosis
-
In the second half of 2024, we plan to initiate a new cohort in the Phase 3 clinical trial, RIDGELINE, evaluating the vanzacaftor triple in children with CF 2 to 5 years of age who have at least one F508del mutation or a mutation responsive to triple combination CFTR modulators.
-
In collaboration with Moderna, Inc. (“Moderna”), we are developing VX-522, a nebulized mRNA therapy for the treatment of people with CF who do not produce full-length CFTR protein. We continue to enroll and dose patients in the multiple ascending dose portion of the Phase 1/2 clinical trial of VX-522 in people with CF. We expect to share data from this clinical trial in late 2024 or early 2025.
Sickle Cell Disease and Transfusion-Dependent Beta Thalassemia
-
We have completed enrollment in two global Phase 3 clinical trials evaluating CASGEVY in children 5 to 11 years of age with SCD or TDT.
-
We continue to work on preclinical assets for myeloablative conditioning agents that would have milder side-effects and could be used in connection with CASGEVY, which could broaden the eligible patient population.
Acute Pain
-
We expect to initiate a Phase 2 clinical trial evaluating the oral formulation of VX-993, a next-generation selective NaV1.8 inhibitor, for the treatment of moderate-to-severe acute pain in 2024.
-
The FDA cleared the investigational new drug application (“IND”) for an intravenous formulation of VX-993 for the treatment of moderate-to-severe acute pain. We have initiated the Phase 1 clinical trial.
-
We are advancing multiple NaV1.7 inhibitors through research and earlier stages of development, which could be used alone or in combination with NaV1.8 inhibitors for the treatment of acute and peripheral neuropathic pain.
Peripheral Neuropathic Pain
-
Following a successful end-of-Phase 2 meeting with the FDA, we expect to initiate the Phase 3 pivotal program evaluating suzetrigine in people with diabetic peripheral neuropathy, a common form of chronic peripheral neuropathic pain, in the second half of 2024. The FDA has granted suzetrigine Breakthrough Therapy designation in this indication.
-
We continue to enroll and dose patients in the Phase 2 clinical trial evaluating suzetrigine in people with lumbosacral radiculopathy, a second type of peripheral neuropathic pain. We expect to complete enrollment in the Phase 2 clinical trial by the end of 2024.
-
We expect to initiate a Phase 2 clinical trial evaluating the oral formulation of VX-993 for the treatment of peripheral neuropathic pain in 2024.
APOL1-Mediated Kidney Disease
-
Inaxaplin is our small molecule for the treatment of APOL1-mediated kidney disease (“AMKD”), including APOL1-mediated focal segmental glomerulosclerosis (“FSGS”). Based on the totality of the unblinded data reviewed by the Independent Data Safety Monitoring Committee (“IDMC”), we have advanced into the Phase 3 portion of the global Phase 2/3 pivotal clinical trial in people with AMKD. The trial has been expanded to include adolescents 10 to 17 years of age with AMKD.
-
The clinical trial is designed to have a pre-planned interim analysis at Week 48 evaluating eGFR slope, supported by a percentage change from baseline in proteinuria, in the inaxaplin arm versus placebo. If positive, we expect that the interim analysis may serve as the basis to seek accelerated approval in the U.S.
Type 1 Diabetes
-
VX-880 is an allogeneic stem cell-derived, fully differentiated, insulin-producing islet cell replacement therapy, using standard immunosuppression to protect the implanted cells. We are evaluating VX-880 as a potential treatment for type 1 diabetes (“T1D”) in a sequential, three-part Phase 1/2 clinical trial. Based on the totality of the data reviewed by the IDMC, we have resumed dosing in the Phase 1/2 clinical trial in people with T1D and impaired awareness of hypoglycemia and recurrent hypoglycemic events. Enrollment in Parts A, B and C of the global 17-patient clinical trial is complete and we expect to complete dosing in the coming months.
-
Our second Phase 1/2 program in T1D evaluates VX-264, which encapsulates the same VX-880 islet cells in a novel device designed to eliminate the need for immunosuppression. This trial is a sequential, multi-part study to evaluate the safety, tolerability and efficacy of VX-264. We have completed Part A of the clinical trial and initiated Part B.
-
Our hypoimmune islet cell program uses CRISPR/Cas9 technology to gene-edit the same allogeneic stem cell-derived, fully differentiated islet cells used in the VX-880 and VX-264 programs. The goal is to cloak the cells from the immune system to explore another possible path to eliminate the need for immunosuppressive therapy. This program continues to progress through the research stage.
Myotonic Dystrophy Type 1
-
Our lead approach for myotonic dystrophy type 1 (“DM1”), VX-670, was in-licensed from Entrada Therapeutics, Inc. (“Entrada”). VX-670 is an oligonucleotide connected to a cyclic peptide to promote effective delivery into cells, which holds the potential to address the underlying cause of DM1.
-
Our IND in the U.S. for VX-670 has cleared, as have the clinical trial applications in Canada, the U.K. and the E.U, and the clinical trial notification in Australia. Enrollment and dosing are underway.
Autosomal Dominant Polycystic Kidney
- We have initiated a Phase 1 clinical trial in healthy volunteers evaluating VX-407, our first-in-class small molecule corrector that targets the underlying cause of autosomal dominant polycystic kidney disease (“ADPKD”) in people with a subset of PKD1 variants.
Investment in External Innovation
-
In April 2024, we entered into an agreement and plan of merger (the “Alpine Merger Agreement”) to acquire all of the issued and outstanding shares of common stock of Alpine Immune Sciences, Inc. (“Alpine”) for approximately $4.9 billion. Pursuant to the Alpine Merger Agreement, we commenced a tender offer to purchase all of the outstanding shares of common stock of Alpine for $65 per share in cash. We expect the transaction to close in the second quarter of 2024, subject to certain customary conditions, including the completion of the tender offer, the expiration of the waiting period under the Hart-Scott-Rodino Antitrust Improvements Act, and other conditions.
-
Alpine’s lead molecule, povetacicept, is a highly potent and effective dual antagonist of BAFF (“B cell activating factor”) and APRIL (“a proliferation-inducing ligand”). Through Phase 2 development, povetacicept has shown potential best-in-class efficacy in IgA nephropathy (“IgAN”), a serious, progressive, autoimmune disease of the kidney that can lead to end-stage-renal disease. There are no approved therapies that target the underlying cause of IgAN, which is the most common cause of primary (idiopathic) glomerulonephritis worldwide, affecting approximately 130,000 people in the U.S. We expect povetacicept will enter Phase 3 clinical development in IgAN in the second half of 2024. Phase 1b/2 clinical trials in autoimmune renal diseases and cytopenias are ongoing with data expected in 2024.
-
We achieved a clinical milestone for VX-670 in DM1 in the first quarter of 2024, resulting in a $75 million milestone payable to Entrada.
Our Business Environment
In the first quarter of 2024, our net product revenues came from the sale of our medicines for the treatment of CF. Our CF strategy involves continuing to develop and obtain approval and reimbursement for treatment regimens that will provide benefits to all people with CF and increasing the number of people with CF eligible and able to receive our medicines, including through label expansions, expanded reimbursement, and the development of new medicines. We are advancing our pipeline of product candidates for the treatment of serious diseases outside of CF, including CASGEVY, which has received marketing approvals in the U.S., the E.U., the U.K., Saudi Arabia, and Bahrain for the treatment of SCD and TDT.
Our strategy is to combine transformative advances in the understanding of causal human biology and the science of therapeutics to discover and develop innovative medicines. This approach includes advancing multiple compounds or therapies from each program, spanning multiple modalities, into early clinical trials to obtain patient data that can inform selection of the most promising therapies for later-stage development, as well as to inform discovery and development efforts. We aim to rapidly follow our first-in-class therapies that achieve proof-of-concept with potential best-in-class candidates to provide durable clinical and commercial success.
In pursuit of new product candidates and therapies in specialty markets, we invest in research and development. We believe that pursuing research in diverse areas allows us to balance the risks inherent in product development and may provide product candidates that will form our pipeline in future years. To supplement our internal research programs, we acquire technologies and programs and collaborate with biopharmaceutical and technology companies, leading academic
research institutions, government laboratories, foundations and other organizations, as needed, to advance research in our areas of therapeutic interest and to access technologies needed to execute on our strategy.
Discovery and development of a new pharmaceutical or biological product is a difficult and lengthy process that requires significant financial resources along with extensive technical and regulatory expertise. Across the industry, most potential drug or biological products never progress into development, and most products that do advance into development never receive marketing approval. Our investments in product candidates are subject to considerable risks. We closely monitor the results of our discovery, research, clinical trials and nonclinical studies and frequently evaluate our product development programs in light of new data and scientific, business and commercial insights, with the objective of balancing risk and potential. This process can result in rapid changes in focus and priorities as new information becomes available and as we gain additional understanding of our ongoing programs and potential new programs, as well as those of our competitors.
Our business also requires ensuring appropriate manufacturing and reimbursement of our products. As we advance our product candidates through clinical development toward commercialization and market and sell our approved products, we build and maintain our supply chain and quality assurance resources. We rely on a global network of third parties, including some in China, and our internal capabilities to manufacture and distribute our products for commercial sale and post-approval clinical trials and to manufacture and distribute our product candidates for clinical trials. In addition to establishing supply chains for each new approved product, we adapt our supply chain for existing products to include additional formulations or to increase scale of production for existing products as needed. Our foreign third-party manufacturers and suppliers may be subject to U.S. legislation, including the BIOSECURE Act, sanctions, trade restrictions and other foreign regulatory requirements which could increase costs or reduce the supply of material available to us, or delay the procurement or supply of such material. The processes for cell and genetic therapies can be more complex than those required for small molecule drugs and require additional investments in different systems, equipment, facilities and expertise. We are focused on ensuring the stability of the supply chains for our current products, as well as for our pipeline programs.
Sales of our products depend, to a large degree, on the extent to which our products are reimbursed by third-party payors, such as government health programs, commercial insurance and managed health care organizations. Reimbursement for our products, including our potential pipeline therapies, cannot be assured and may take significant periods of time to obtain. We dedicate substantial management and other resources to obtain and maintain appropriate levels of reimbursement for our products from third-party payors, including governmental organizations in the U.S. and ex-U.S. markets.
In the U.S., we have worked successfully with third-party payors to promptly obtain appropriate levels of reimbursement for our CF medicines. In addition, we are working with U.S. government and commercial payors with respect to CASGEVY. We anticipate broad access with government and commercial payors for CASGEVY in the U.S., and we have recently entered into multiple agreements with government and commercial health insurance providers to provide access to CASGEVY. We plan to continue to engage in discussions with numerous commercial insurers and managed health care organizations, along with government health programs that are typically managed by authorities in the individual states, to ensure that payors recognize the significant benefits that all of our therapies provide and provide patients with appropriate levels of access to our medicines and therapies now and in the future. We cannot, however, predict how recent changes in the law, including through the Inflation Reduction Act of 2022 and passage of state laws (e.g., transparency laws and prescription drug affordability boards), will affect our ability to negotiate successfully with third-party payors and distribute our products. Similarly, in ex-U.S. markets, we seek government reimbursement for our medicines on a country-by-country or region-by-region basis, as required. This is necessary for each new medicine, as well as for label expansions for our current medicines. We are working with ex-U.S. payors with respect to CASGEVY, and we are pursuing long-term reimbursement agreements. We have secured reimbursed access for people with SCD or TDT in Saudi Arabia and Bahrain and for people with TDT in France through an expanded access program. We expect to continue to focus significant resources to expand and maintain reimbursement for our CF medicines, CASGEVY and, ultimately, pipeline therapies, in U.S. and ex-U.S. markets.
Strategic Transactions
Acquisitions
As part of our business strategy, we seek to acquire technologies, products, product candidates and other businesses that are aligned with our corporate and research and development strategies and complement and advance our ongoing research and development efforts. We have engaged in a number of acquisitions of privately held biotechnology companies over the last several years. In 2019, we acquired Semma Therapeutics, Inc., pursuant to which we established our T1D program, and Exonics Therapeutics, Inc., which enhanced our gene-editing capabilities to repair mutations that cause severe neuromuscular diseases, such as DM1 and Duchenne muscular dystrophy. In 2022, we acquired ViaCyte, Inc. (“ViaCyte”), which had intellectual property, tools, technologies and assets with the potential to accelerate development of our T1D programs. Our
acquisition of ViaCyte, for $315.0 million, was accounted for as a business combination. As of the acquisition date, the cash payment was allocated primarily to goodwill and the fair value of an in-process research and development asset.
In 2023 and 2022, we also acquired programs that were accounted for as asset acquisitions resulting in $47.5 million and $60.0 million of AIPR&D, respectively.
In April 2024, we entered into the Alpine Merger Agreement to acquire Alpine as described above. Alpine is a publicly traded biotechnology company focused on discovering and developing innovative, protein-based immunotherapies. Alpine’s lead molecule, povetacicept, has shown potential best-in-class efficacy in IgAN through Phase 2 development. We will pay approximately $4.9 billion in cash when the transaction closes and intend to fund the acquisition with our cash and cash equivalents. We will account for the acquisition in the period that it closes.
Collaboration and In-Licensing Arrangements
We enter into arrangements with third parties, including collaboration and licensing arrangements, for the development, manufacture and commercialization of products, product candidates and other technologies that have the potential to complement our ongoing research and development efforts. Over the last several years, we entered into collaboration agreements with a number of companies, including CRISPR Therapeutics AG (“CRISPR”), Entrada, and Moderna. Generally, when we in-license a technology or product candidate, we make upfront payments to the collaborator, assume the costs of the program and/or agree to make contingent payments, which could consist of milestone, royalty and option payments. Most of these collaboration payments are expensed as AIPR&D, including a $75.0 million milestone due to Entrada in the first quarter of 2024, and our upfront payments of $225.1 million to Entrada and $100.0 million to CRISPR related to T1D in the first quarter of 2023. These payments were expensed to AIPR&D because they were primarily attributable to acquired in-process research and development for which there was no alternative future use. However, depending on many factors, including the structure of the collaboration, the stage of development of the acquired technology, the significance of the in-licensed product candidate to the collaborator’s operations and the other activities in which our collaborators are engaged, the accounting for these transactions can vary significantly. We expect to continue to identify and evaluate collaboration and licensing opportunities that may be similar to or different from the collaborations and licenses that we have engaged in previously.
Acquired In-Process Research and Development Expenses
In the first quarter of 2024 and 2023, our AIPR&D included $76.8 million and $347.1 million, respectively, related to upfront, contingent milestone, or other payments pursuant to our business development transactions, including the collaborations, licenses of third-party technologies, and asset acquisitions described above. Please refer to Note C, “Collaboration, License and Other Arrangements,” for further information regarding our collaboration, in-license agreements and asset acquisitions.
RESULTS OF OPERATIONS
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2024 | 2023 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages and per share amounts) | |||||||||||||||||||||||||||||||||||
| Product revenues, net | $ | 2,690.6 | $ | 2,374.8 | 13% | ||||||||||||||||||||||||||||||
| Operating costs and expenses | 1,551.1 | 1,595.8 | (3)% | ||||||||||||||||||||||||||||||||
| Income from operations | 1,139.5 | 779.0 | 46% | ||||||||||||||||||||||||||||||||
| Other non-operating income, net | 139.6 | 112.5 | 24% | ||||||||||||||||||||||||||||||||
| Provision for income taxes | 179.5 | 191.7 | (6)% | ||||||||||||||||||||||||||||||||
| Net income | $ | 1,099.6 | $ | 699.8 | 57% | ||||||||||||||||||||||||||||||
| Net income per diluted common share | $ | 4.21 | $ | 2.69 | |||||||||||||||||||||||||||||||
| Diluted shares used in per share calculations | 261.1 | 260.3 | |||||||||||||||||||||||||||||||||
Product Revenues, net
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2024 | 2023 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| TRIKAFTA/KAFTRIO | $ | 2,483.6 | $ | 2,096.7 | 18% | ||||||||||||||||||||||||||||||
| Other CF products | 207.0 | 278.1 | (26)% | ||||||||||||||||||||||||||||||||
| Product revenues, net | $ | 2,690.6 | $ | 2,374.8 | 13% | ||||||||||||||||||||||||||||||
In the first quarter of 2024, our net product revenues increased by $315.8 million, or 13%, as compared to the first quarter of 2023. The increase was primarily due to the continued strong uptake of TRIKAFTA/KAFTRIO in ex-U.S. markets and label extensions in younger age groups and performance of TRIKAFTA in the U.S., following the launch of TRIKAFTA in children with CF 2 to 5 years of age. Decreases in revenues for our CF products other than TRIKAFTA/KAFTRIO were primarily the result of patients switching from these medicines to TRIKAFTA/KAFTRIO.
Our net product revenues from the U.S. and from ex-U.S. markets were as follows:
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2024 | 2023 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| United States | $ | 1,519.9 | $ | 1,403.8 | 8% | ||||||||||||||||||||||||||||||
| ex-U.S. | 1,170.7 | 971.0 | 21% | ||||||||||||||||||||||||||||||||
| Product revenues, net | $ | 2,690.6 | $ | 2,374.8 | 13% |
Operating Costs and Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2024 | 2023 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Cost of sales | $ | 342.6 | $ | 266.9 | 28% | ||||||||||||||||||||||||||||||
| Research and development expenses | 789.1 | 742.6 | 6% | ||||||||||||||||||||||||||||||||
| Acquired in-process research and development expenses | 76.8 | 347.1 | (78)% | ||||||||||||||||||||||||||||||||
| Selling, general and administrative expenses | 342.7 | 241.1 | 42% | ||||||||||||||||||||||||||||||||
| Change in fair value of contingent consideration | (0.1) | (1.9) | ** | ||||||||||||||||||||||||||||||||
| Total costs and expenses | $ | 1,551.1 | $ | 1,595.8 | (3)% | ||||||||||||||||||||||||||||||
| ** Not meaningful |
Cost of Sales
Our cost of sales primarily consists of third-party royalties payable on net sales of our CF products as well as the cost of producing inventories. Pursuant to our agreement with the Cystic Fibrosis Foundation, our tiered third-party royalties on sales of TRIKAFTA/KAFTRIO, SYMDEKO/SYMKEVI, KALYDECO, and ORKAMBI, calculated as a percentage of net sales, range from the single digits to the sub-teens, with lower royalties on sales of TRIKAFTA/KAFTRIO than for our other products. Over the last several years, our cost of sales has been increasing due to increased net product revenues. Our cost of sales as a percentage of our net product revenues increased to 13% in the first quarter of 2024 as compared to 11% in the first quarter of 2023, primarily due to cost of sales associated with CASGEVY following its regulatory approval in the fourth quarter of 2023.
Research and Development Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2024 | 2023 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Research expenses | $ | 196.1 | $ | 166.8 | 18% | ||||||||||||||||||||||||||||||
| Development expenses | 593.0 | 575.8 | 3% | ||||||||||||||||||||||||||||||||
| Total research and development expenses | $ | 789.1 | $ | 742.6 | 6% |
Our research and development expenses include internal and external costs incurred for research and development of our products and product candidates. We do not assign our internal costs, such as salary and benefits, stock-based compensation expense, laboratory supplies and other direct expenses and infrastructure costs, to individual products or product candidates, because the employees within our research and development groups typically are deployed across multiple research and development programs. We assign external costs of services provided to us by clinical research organizations and other outsourced research by individual program. Our internal costs are greater than our external costs. All research and development costs for our products and product candidates are expensed as incurred.
Since January 2022, we have incurred approximately $7.2 billion in total research and development and AIPR&D expenses associated with product discovery and development. The successful development of our product candidates is highly uncertain and subject to a number of risks. In addition, the duration of clinical trials may vary substantially according to the type, complexity and novelty of the product candidate and the disease indication being targeted. The FDA and comparable agencies in foreign countries impose substantial requirements on the introduction of therapeutic pharmaceutical products, typically requiring lengthy and detailed laboratory and clinical testing procedures, sampling activities and other costly and time-consuming procedures. Data obtained from nonclinical and clinical activities at any step in the testing process may be adverse and lead to discontinuation or redirection of development activities. Data obtained from these activities also are susceptible to varying interpretations, which could delay, limit or prevent regulatory approval. The duration and cost of discovery, nonclinical studies and clinical trials may vary significantly over the life of a project and are difficult to predict. Therefore, accurate and meaningful estimates of the ultimate costs to bring our product candidates to market are not available.
Any estimates regarding development and regulatory timelines for our product candidates are highly subjective and subject to change. Until we have data from Phase 3 clinical trials, we cannot make a meaningful estimate regarding when, or if, a clinical development program will generate revenues and cash flows.
Research Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2024 | 2023 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Research Expenses: | |||||||||||||||||||||||||||||||||||
| Salary and benefits | $ | 53.0 | $ | 45.5 | 16% | ||||||||||||||||||||||||||||||
| Stock-based compensation expense | 29.9 | 20.2 | 48% | ||||||||||||||||||||||||||||||||
| Outsourced services and other direct expenses | 64.4 | 53.6 | 20% | ||||||||||||||||||||||||||||||||
| Infrastructure costs | 48.8 | 47.5 | 3% | ||||||||||||||||||||||||||||||||
| Total research expenses | $ | 196.1 | $ | 166.8 | 18% | ||||||||||||||||||||||||||||||
Our research expenses have been increasing over the last several years as we have invested in our pipeline and expanded our cell and genetic therapy capabilities, resulting in increased headcount, and outside services and other direct expenses. We expect to continue to invest in our research programs with a focus on creating transformative medicines for serious diseases.
Development Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2024 | 2023 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Development Expenses: | |||||||||||||||||||||||||||||||||||
| Salary and benefits | $ | 170.1 | $ | 144.2 | 18% | ||||||||||||||||||||||||||||||
| Stock-based compensation expense | 89.5 | 56.1 | 60% | ||||||||||||||||||||||||||||||||
| Outsourced services and other direct expenses | 236.1 | 295.3 | (20)% | ||||||||||||||||||||||||||||||||
| Infrastructure costs | 97.3 | 80.2 | 21% | ||||||||||||||||||||||||||||||||
| Total development expenses | $ | 593.0 | $ | 575.8 | 3% | ||||||||||||||||||||||||||||||
Our development expenses increased by $17.2 million, or 3%, in the three months ended March 31, 2024 as compared to the three months ended March 31, 2023, primarily due to increased headcount costs, including stock-based compensation expense, and infrastructure costs to support our clinical trials, partially offset by decreased outsourced services and other direct expenses.
We are investing significantly in internal headcount and in infrastructure to support our advancing pipeline. Additional headcount over the last several years has resulted in increased stock-based compensation expense. Our stock-based compensation expense has historically fluctuated, and is expected to continue to fluctuate from one period to another based on the probability of achieving milestones associated with our performance-based awards. Our outsourced services and other direct expenses decreased as compared to the three months ended March 31, 2023 primarily due to lower clinical trial costs resulting from the commercialization of CASGEVY, as well as advancements in both our suzetrigine program in acute pain and the vanzacaftor triple for CF. These decreased clinical trial expenses were partially offset by increased costs to support our T1D program.
Acquired In-process Research and Development Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2024 | 2023 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Acquired in-process research and development expenses | $ | 76.8 | $ | 347.1 | (78)% | ||||||||||||||||||||||||||||||
AIPR&D in the first quarter of 2024 was primarily related to the $75.0 million milestone due to Entrada. AIPR&D in the first quarter of 2023 was primarily related to our upfront payments of $225.1 million to Entrada and $100.0 million to CRISPR. Our AIPR&D has historically fluctuated, and is expected to continue to fluctuate, from one period to another due to upfront, contingent milestone, and other payments pursuant to our existing and future business development transactions, including collaborations, licenses of third-party technologies, and asset acquisitions.
Selling, General and Administrative Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2024 | 2023 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Selling, general and administrative expenses | $ | 342.7 | $ | 241.1 | 42% |
Selling, general and administrative expenses increased by 42% in the first quarter of 2024 as compared to the first quarter of 2023, primarily due to increased investments to support the launches of our therapies globally and prepare for near-term launches of multiple potential new products.
Contingent Consideration
The fair value of our contingent consideration decreased by $0.1 million and $1.9 million in the first quarter of 2024 and 2023, respectively.
Other Non-Operating Income (Expense), Net
Interest Income
Interest income increased to $181.2 million in the first quarter of 2024, as compared to $122.6 million in the first quarter of 2023, primarily due to increased market interest rates, increased cash equivalents and available-for-sale debt securities and mix of available-for-sale debt securities earning higher yields. Our future interest income is dependent on the amount of, and prevailing market interest rates on, our outstanding cash equivalents and available-for-sale debt securities.
Interest Expense
Interest expense was $10.4 million and $11.4 million in the first quarter of 2024 and 2023, respectively. The majority of our interest expense in these periods was related to imputed interest expense associated with our leased corporate headquarters in Boston.
Other Income (Expense), Net
Other income (expense), net was expense of $31.2 million and income of $1.3 million in the first quarter of 2024 and 2023, respectively. These amounts related primarily to net unrealized gains or losses resulting from changes in the fair value of our strategic equity investments, which consist of investments in our collaborators that may be public or private companies. To the extent that we continue to hold strategic equity investments in publicly traded companies, we expect that due to the volatility of the stock price of biotechnology companies, our other income (expense), net will fluctuate in future periods based on increases or decreases in the fair value of our strategic equity investments. As of March 31, 2024, the fair value of our investments in publicly traded companies was $43.4 million.
Income Taxes
Our effective tax rate fluctuates from period to period due to the global nature of our operations. The factors that most significantly impact our effective tax rate include changes in tax laws, variability in the allocation of our taxable earnings among multiple jurisdictions, the amount and characterization of our research and development expenses, the levels of certain deductions and credits, adjustments to the value of our uncertain tax positions, acquisitions and third-party collaboration and licensing transactions.
We recorded provisions for income taxes of $179.5 million and $191.7 million in the first quarter of 2024 and 2023, respectively. Our effective tax rate of 14.0% in the first quarter of 2024 was lower than the U.S. statutory rate primarily due to changes in our unrecognized tax positions as well as excess tax benefits related to stock-based compensation. Our effective tax rate of 21.5% in the first quarter of 2023 was higher than the U.S. statutory rate primarily due to an increase in our unrecognized tax positions partially offset by excess tax benefits related to stock-based compensation.
LIQUIDITY AND CAPITAL RESOURCES
The following table summarizes the components of our financial condition as of March 31, 2024 and December 31, 2023:
| As of March 31, 2024 | As of December 31, 2023 | Change | |||||||||||||||
| (in millions, except percentages) | |||||||||||||||||
| Cash, cash equivalents and marketable securities: | |||||||||||||||||
| Cash and cash equivalents | $ | 9,158.0 | $ | 10,369.1 | |||||||||||||
| Marketable securities | 1,013.3 | 849.2 | |||||||||||||||
| Long-term marketable securities | 4,381.4 | 2,497.8 | |||||||||||||||
| Total cash, cash equivalents and marketable securities | $ | 14,552.7 | $ | 13,716.1 | 6% | ||||||||||||
| Working Capital: | |||||||||||||||||
| Total current assets | $ | 13,288.7 | $ | 14,144.2 | (6)% | ||||||||||||
| Total current liabilities | (3,795.9) | (3,547.4) | 7% | ||||||||||||||
| Total working capital | $ | 9,492.8 | $ | 10,596.8 | (10)% |
Working Capital
As of March 31, 2024, total working capital was $9.5 billion, which represented a decrease of $1.1 billion from $10.6 billion as of December 31, 2023. The decrease in total working capital during the first quarter of 2024 was primarily due to net purchases of long-term marketable securities of $2.1 billion.
Cash Flows
| Three Months Ended March 31, | |||||||||||
| 2024 | 2023 | ||||||||||
| (in millions) | |||||||||||
| Net cash provided by (used in): | |||||||||||
| Operating activities | $ | 1,306.6 | $ | 899.9 | |||||||
| Investing activities | $ | (2,136.4) | $ | (1,833.6) | |||||||
| Financing activities | $ | (357.5) | $ | (294.7) |
Operating Activities
Cash provided by operating activities was $1.3 billion in the first quarter of 2024 as compared to $899.9 million in the first quarter of 2023, primarily due to increased net income resulting from increased net product revenues.
Investing Activities
Cash used in investing activities were $2.1 billion and $1.8 billion in the first quarter of 2024 and 2023, respectively, which primarily relate to net purchases of available-for-sale debt securities.
Financing Activities
Cash used in financing activities were $357.5 million and $294.7 million in the first quarter of 2024 and 2023, respectively. The largest portions of our financing activities in each of these periods were payments related to our employee stock benefit plans and repurchases of our common stock pursuant to our share repurchase program.
Sources and Uses of Liquidity
We intend to rely on our existing cash, cash equivalents and current marketable securities together with cash flows from product sales as our primary source of liquidity. We expect that cash flows from our product sales together with our cash, cash equivalents and current marketable securities will be sufficient to fund our operations for at least the next twelve months. The adequacy of our available funds to meet our future operating and capital requirements will depend on many factors, including our future product sales, and the potential introduction of one or more of our other product candidates to the market, our business development activities, and the number, breadth, cost and prospects of our research and development programs.
Credit Facilities & Financing Strategy
We may borrow up to a total of $500.0 million pursuant to a revolving credit facility that we entered into in July 2022 and could repay and reborrow amounts under this revolving credit agreement without penalty. Subject to certain conditions, we could request that the borrowing capacity be increased by an additional $500.0 million, for a total of $1.0 billion. Negative covenants in our credit agreement could prohibit or limit our ability to access this source of liquidity. As of March 31, 2024, the facility was undrawn, and we were in compliance with these covenants.
We may also raise additional capital by borrowing under credit agreements, through public offerings or private placements of our securities or securing new collaborative agreements or other methods of financing. We will continue to manage our capital structure and will consider all financing opportunities, whenever they may occur, that could strengthen our long-term liquidity profile. There can be no assurance that any such financing opportunities will be available on acceptable terms, if at all.
Future Capital Requirements
We have significant future capital requirements, including:
-
In April 2024, we entered into the Alpine Merger Agreement to acquire Alpine for approximately $4.9 billion in cash, as described above. The transaction is expected to close in the second quarter of 2024, subject to certain customary closing conditions. We intend to fund the acquisition with our cash and cash equivalents.
-
Expected operating expenses to conduct research and development activities, manufacture and commercialize our existing and future products, and to operate our organization.
-
Cash that we pay for income taxes.
-
Royalties we pay related to sales of our CF products.
-
Facility, operating and finance lease obligations.
-
Firm purchase obligations related to our supply and manufacturing processes.
In addition, other potential significant future capital requirements may include:
-
We have entered into certain business development-related and strategic agreements with third parties that include the funding of certain research, development, manufacturing and commercialization efforts. Certain of our transactions, including collaborations, licensing arrangements, and asset acquisitions, include the potential for future milestone and royalty payments by us upon the achievement of pre-established developmental and regulatory targets and/or commercial targets. Other transactions include the potential for future lease-related expenses and other costs. Our obligation to fund these research and development and commercialization efforts and to pay these potential milestones, expenses and royalties is contingent upon continued involvement in the programs and/or the lack of any adverse events that could cause their discontinuance. We may enter into additional business development transactions and strategic agreements, including acquisitions, collaborations, licensing arrangements and equity investments, which require additional capital.
-
To the extent we borrow amounts under our existing credit agreement, we would be required to repay any outstanding principal amounts in 2027.
-
As of March 31, 2024, we had $2.4 billion remaining authorization available under our Share Repurchase Program that our Board of Directors approved in February 2023. We expect to fund repurchases of our common stock through a combination of cash on hand and cash generated by operations. This program does not have an expiration date and can be discontinued at any time.
Other than our potential payment to acquire Alpine noted above, there have not been any material changes to our future capital requirements disclosed in our Annual Report on Form 10-K for the year ended December 31, 2023, which was filed with the Securities and Exchange Commission, or SEC, on February 15, 2024.
CRITICAL ACCOUNTING POLICIES AND ESTIMATES
Our discussion and analysis of our financial condition and results of operations are based upon our condensed consolidated financial statements prepared in accordance with generally accepted accounting principles in the U.S. The preparation of these financial statements requires us to make certain estimates and assumptions that affect the reported amounts of assets and liabilities, the disclosure of contingent assets and liabilities at the date of the condensed consolidated financial statements and the reported amounts of revenues and expenses during the reported periods. These items are monitored and analyzed by management for changes in facts and circumstances, and material changes in these estimates could occur in the future. Changes in estimates are reflected in reported results for the period in which the change occurs. We base our estimates on historical experience and various other assumptions that we believe to be reasonable under the circumstances. Actual results may differ from our estimates if past experience or other assumptions do not turn out to be substantially accurate. During the three months ended March 31, 2024, there were no material changes to our critical accounting policies as reported in our Annual Report on Form 10-K for the year ended December 31, 2023, which was filed with the SEC on February 15, 2024.
RECENT ACCOUNTING PRONOUNCEMENTS
For a discussion of recent accounting pronouncements, please refer to Note A, “Basis of Presentation and Accounting Policies.”
Previous: Item 1. Financial Statements · Next: Item 3. Quantitative and Qualitative Disclosures About Market Risk