 filings.tools
Item 2. Management’s Discussion and Analysis of Financial Condition and Results of Operations
64K characters. Original on sec.gov · Markdown
Item 2. Management’s Discussion and Analysis of Financial Condition and Results of Operations
OVERVIEW
We are a global biotechnology company that invests in scientific innovation to create transformative medicines for people with serious diseases, with a focus on specialty markets. We have seven approved medicines: five that treat the underlying cause of cystic fibrosis (“CF”), a life-threatening genetic disease, one that treats severe sickle cell disease (“SCD”) and transfusion dependent beta thalassemia (“TDT”), life shortening inherited blood disorders, and one that treats moderate-to-severe acute pain. Our clinical-stage pipeline includes programs in CF, SCD, beta thalassemia, acute and peripheral neuropathic pain, APOL1-mediated kidney disease, IgA nephropathy and other autoimmune renal diseases and cytopenias, type 1 diabetes, myotonic dystrophy type 1, and autosomal dominant polycystic kidney disease.
In December 2024, the U.S. Food and Drug Administration (the “FDA”) approved ALYFTREK (vanzacaftor/tezacaftor/deutivacaftor), our once-daily next-in-class triple combination for the treatment of people with CF 6 years of age and older, and our fifth CF medicine. ALYFTREK is also approved in the United Kingdom (the “U.K.”). Collectively, our five medicines, led by TRIKAFTA/KAFTRIO (elexacaftor/tezacaftor/ivacaftor and ivacaftor), are being used to treat nearly three quarters of the approximately 94,000 people with CF in the U.S., Europe, Australia, and Canada. Through approvals of new medicines, label expansions, and expanded reimbursement, we are focused on increasing the number of people with CF who are eligible and able to receive our medicines. In addition, we are evaluating our CF medicines in additional patient populations, including younger children, with the goal of having small molecule treatments for all people who have at least one mutation in their CFTR gene that is responsive to our CFTR modulators. We also are pursuing messenger ribonucleic acid (“mRNA”) and genetic therapies for people with CF who do not make full-length CFTR protein and, as a result, cannot benefit from our current CF medicines.
CASGEVY (exagamglogene autotemcel), our ex-vivo, non-viral CRISPR/Cas9 gene-edited cell therapy, is approved in the U.S., the European Union (“E.U.”), the U.K., the Kingdom of Saudi Arabia (“Saudi Arabia”), the Kingdom of Bahrain (“Bahrain”), the United Arab Emirates (the “UAE”), Switzerland and Canada for the treatment of people 12 years of age and older with SCD or TDT. We estimate approximately 60,000 people with severe SCD or TDT are or could become eligible for CASGEVY in the U.S., Canada, Europe, Saudi Arabia and Bahrain.
In January 2025, the FDA approved JOURNAVX, our selective non-opioid NaV1.8 pain signal inhibitor, for the treatment of people with moderate-to-severe acute pain. We have begun our commercial launch of JOURNAVX in the U.S. for eligible adults. In addition, we are enrolling and dosing patients in a Phase 3 clinical trial evaluating suzetrigine for the treatment of diabetic peripheral neuropathy, a common form of peripheral neuropathic pain.
Financial Highlights
| Revenues | In the first quarter of 2025, our net product revenues increased to $2.8 billion as compared to $2.7 billion in the first quarter of 2024, primarily due to the continued performance of TRIKAFTA/KAFTRIO and initial contributions from ALYFTREK. | ||||
| Expenses | Our total research and development (“R&D”), acquired in-process research and development (“AIPR&D”), and selling, general and administrative (“SG&A”) expenses increased to $1.4 billion in the first quarter of 2025 as compared to $1.2 billion in the first quarter of 2024, primarily due to increased commercial investments to support the launch of JOURNAVX and continued investment to support additional therapies in mid-to-late stage development, partially offset by decreased AIPR&D. Cost of sales was 13% in each of the first quarter of 2025 and 2024. | ||||
| Cash | Our total cash, cash equivalents and marketable securities increased to $11.4 billion as of March 31, 2025 as compared to $11.2 billion as of December 31, 2024 primarily due to cash flows provided by our operating activities partially offset by repurchases of our common stock. |
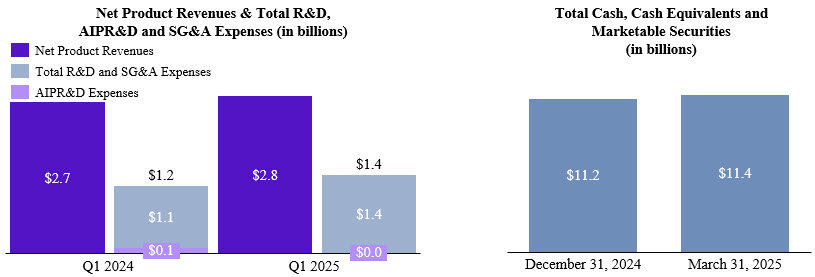
Note: Charts above may not add due to rounding.
Business Updates
Marketed Products
Cystic Fibrosis
We expect to grow our CF business by increasing the number of people with CF who are eligible and able to receive our medicines.
Recent and anticipated progress in activities expanding our CF business is included below:
-
ALYFTREK, the once-daily next-in-class combination CFTR modulator for the treatment of people with CF 6 years of age and older who have at least one F508del mutation or another responsive mutation in the CFTR gene, is approved in the U.S. and the U.K. We are working with the National Health Service in the U.K. to secure coverage for eligible patients.
-
The European Medical Agency’s (the “EMA”) Committee for Medicinal Products for Human Use adopted a positive opinion for ALYFTREK for the treatment of people with CF 6 years of age and older who have at least one non-class I mutation in the CFTR gene. European Commission approval for ALYFTREK is expected in the second half of 2025. Regulatory submissions for ALYFTREK are under review in Canada, Switzerland, Australia and New Zealand.
-
KAFTRIO is approved by the European Commission for the treatment of people with CF 2 years of age and older who have at least one non-class I mutation in the CFTR gene. With this approval, approximately 4,000 people in the European Union are newly eligible for a medicine that treats the underlying cause of their disease. We are working to ensure access for all eligible patients.
Sickle Cell Disease and Beta Thalassemia
-
We entered into a reimbursement agreement with NHS England for eligible people with SCD to access CASGEVY, and a similar agreement in Wales for eligible people with SCD or TDT. Following a positive assessment, we also have finalized national reimbursement in Austria. In the Middle East, following regulatory approval in the UAE, we have entered into a reimbursement agreement for eligible people with SCD or TDT in the majority of emirates.
-
We have activated more than 65 authorized treatment centers globally, and approximately 90 patients have had their first cell collection.
-
We filed a manufacturing license submission with the FDA and we expect to begin manufacturing CASGEVY in Portsmouth, NH in the second half of 2025. This submission is part of the planned ramp of CASGEVY manufacturing capacity as demand for the therapy increases.
Acute Pain
-
In January 2025, the FDA approved JOURNAVX, for the treatment of adults with moderate-to-severe acute pain. JOURNAVX is available and stocked at pharmacies across the U.S., including major national and regional retail pharmacy chains.
-
Since JOURNAVX became available in early March, more than 20,000 prescriptions have been written and filled across the hospital and retail settings in different acute pain conditions, consistent with its broad label.
-
Across commercial and government payers, approximately 94 million lives have covered access to JOURNAVX, and approximately 42 million have unrestricted access (i.e., without the need for prior authorization or step edits). We have reached a formal coverage agreement with a large national Pharmacy Benefit Manager to make JOURNAVX available to their customers, representing approximately 22 million commercial lives. A total of ten state Medicaid plans are also providing unrestricted access to JOURNAVX and 20 more are currently evaluating their policies.
-
More than 50 large healthcare systems have taken steps to initiate pharmacy and therapeutics (“P&T”) committee reviews of JOURNAVX, and some have already added it on formularies.
-
We recently initiated two Phase 4 clinical trials in various moderate-to-severe acute pain conditions to provide additional data on the effectiveness and safety of JOURNAVX as part of real-world clinical practice, in both inpatient and outpatient settings.
-
We expect JOURNAVX to be added to the list of treatments eligible for an add-on payment under the Non-Opioids Prevent Addiction in the Nation (“NOPAIN”) Act, which became effective on January 1, 2025.
Pipeline
We continue to advance a diversified pipeline of potentially transformative medicines for serious diseases utilizing a range of modalities. Recent and anticipated progress in activities supporting these efforts is included below:
Cystic Fibrosis
-
We are enrolling and dosing patients in a Phase 3 clinical trial of TRIKAFTA/KAFTRIO in children 1 to 2 years of age and in a Phase 3 clinical trial of ALYFTREK in children 2 to 5 years of age to expand the labels and enable earlier treatment of children with CF.
-
We continue to advance new oral small molecule combination therapies through preclinical and clinical development. We expect to advance the once-daily, next-generation 3.0 VX-828 combination into a clinical trial in people with CF in 2025.
-
In collaboration with Moderna, Inc. (“Moderna”), we are developing VX-522, a nebulized CFTR mRNA therapy for the treatment of people with CF who do not produce full-length CFTR protein. We have implemented a temporary pause to the multiple ascending dose portion of the Phase 1/2 clinical trial of VX-522 in order to assess a tolerability issue.
Sickle Cell Disease and Transfusion-Dependent Beta Thalassemia
-
We have completed enrollment in two global Phase 3 clinical trials evaluating CASGEVY in children 5 to 11 years of age with SCD or TDT.
-
We continue to advance preclinical assets for myeloablative conditioning agents that would have milder side-effects and could be used in connection with CASGEVY, which could broaden the eligible patient population.
Acute Pain
-
We expect to complete the Phase 2 clinical trial evaluating an oral formulation of VX-993 in the second quarter of 2025. VX-993 is a next-generation selective NaV1.8 pain signal inhibitor, for the treatment of moderate-to-severe acute pain following bunionectomy surgery. We expect to report data from this clinical trial in the second half of 2025.
-
We have completed a Phase 1 clinical trial evaluating an intravenous formulation of VX-993 in healthy volunteers.
-
The FDA has granted Fast Track Designation to VX-993 in moderate-to-severe acute pain in both the oral and intravenous formulations.
Peripheral Neuropathic Pain
-
We are enrolling and dosing people with diabetic peripheral neuropathy, a common form of chronic peripheral neuropathic pain, in a Phase 3 pivotal trial evaluating suzetrigine. The FDA has granted suzetrigine Fast Track designation in peripheral neuropathic pain and Breakthrough Therapy designation in diabetic peripheral neuropathy.
-
We are enrolling and dosing patients in a Phase 2 clinical trial evaluating the oral formulation of VX-993 for the treatment of diabetic peripheral neuropathy.
Type 1 Diabetes
-
Zimislecel is an allogeneic, stem cell-derived, fully differentiated, insulin-producing islet cell replacement therapy, using standard immunosuppression to protect the implanted cells. We are enrolling and dosing patients in the Phase 3 portion of the Phase 1/2/3 clinical trial of zimislecel in people with type 1 diabetes (“T1D”) with severe hypoglycemic events and impaired awareness of hypoglycemia in the U.S., Canada, U.K. and E.U. We expect to complete enrollment and dosing in this pivotal clinical trial in the second quarter of 2025 and we expect to submit marketing applications to global regulators in 2026. We also expect to share updated clinical trial data in the second quarter of 2025.
-
Zimislecel has been granted Regenerative Medicine Advanced Therapy and Fast Track designations from the FDA, Priority Medicines designation from the EMA, and has secured an Innovation Passport under the Innovative Licensing and Access Pathway from the UK Medicines and Healthcare products Regulatory Agency.
-
In March 2025, we announced results from the Phase 1/2 clinical trial evaluating VX-264, which encapsulated zimislecel in an immunoprotective device. VX-264 was generally safe and well-tolerated but did not meet its efficacy endpoint, and we have discontinued development of this program.
-
We are pursuing research-stage programs to evaluate additional approaches that could provide transformative benefit to people with T1D and reduce or eliminate the need for standard immunosuppressive regimens. These approaches include alternative immunosuppressives, gene editing, and novel immunoprotection to encapsulate the islet cells.
IgA Nephropathy, Primary Membranous Nephropathy and Other B Cell-Mediated Diseases
-
We are developing povetacicept, a dual antagonist of B cell activating factor (“BAFF”) and a proliferation-inducing ligand (“APRIL”) cytokines, as a potentially best-in-class approach to treat immunoglobulin A nephropathy (“IgAN”) and primary membranous nephropathy (“pMN”).
-
The global Phase 3 RAINIER trial evaluating povetacicept in people with IgAN has completed enrollment of the interim analysis cohort. The interim analysis will be conducted once this cohort reaches 36 weeks of treatment, with the potential to file for Accelerated Approval in the U.S. in the first half of 2026, if results are supportive. Clinical trials to support the launch of povetacicept for home administration are underway.
-
Based on positive results of povetacicept in pMN in the RUBY-3 clinical trial, we have reached agreement with the FDA to advance povetacicept into pivotal development for this disease. We expect to initiate a single, adaptive Phase 2/3 clinical trial of povetacicept vs. standard-of-care in 2025, and the selected dose will be advanced to Phase 3. We expect to present updated data from the IgAN and pMN cohorts of the RUBY-3 clinical trial at upcoming medical congresses.
-
We are studying additional renal diseases in the RUBY-3 basket trial and hematologic conditions in the RUBY-4 basket trial. We expect to share data and next steps from these trials in 2025.
APOL1-Mediated Kidney Disease
-
Inaxaplin is our small molecule for the treatment of APOL1-mediated kidney disease (“AMKD”). We expect to complete enrollment in the interim analysis cohort of the Phase 3 portion of the global Phase 2/3 pivotal clinical trial (“AMPLITUDE”) in the second half of 2025. We expect to conduct the pre-planned interim analysis once this cohort has been treated for 48 weeks, with potential to file for accelerated approval in the U.S. if the results are supportive.
-
We continue to enroll and dose in the Phase 2 proof-of-concept clinical trial (“AMPLIFIED”) evaluating inaxaplin in people with AMKD and diabetes or other co-morbidities.
Myotonic Dystrophy Type 1
-
Our lead approach for myotonic dystrophy type 1 (“DM1”), VX-670, was in-licensed from Entrada Therapeutics, Inc. (“Entrada”). VX-670 is an oligonucleotide connected to a cyclic peptide to promote effective delivery into cells, which holds the potential to address the underlying cause of DM1.
-
We are enrolling and dosing in the multiple ascending dose portion of the global Phase 1/2 clinical trial evaluating VX-670 in people with DM1, which will evaluate both safety and efficacy.
Autosomal Dominant Polycystic Kidney Disease
-
We have completed a Phase 1 clinical trial in healthy volunteers evaluating VX-407, our first-in-class small molecule corrector that targets the underlying cause of autosomal dominant polycystic kidney disease (“ADPKD”) in people with a subset of PKD1 variants.
-
We expect to advance VX-407 into a Phase 2 proof-of-concept clinical trial in 2025 in people with ADPKD with a subset of variants in the PKD1 gene.
Our Business Environment
In the first quarter of 2025, our net product revenues came primarily from the sale of our medicines for the treatment of CF. Our CF strategy involves continuing to develop and obtain approval and reimbursement for treatment regimens that will provide benefits to all people with CF and increasing the number of people with CF eligible and able to receive our medicines. We are continuing to progress commercialization of CASGEVY, which has received marketing approvals in the U.S., the E.U., the U.K., Saudi Arabia, Bahrain, the UAE, Switzerland and Canada for the treatment of SCD and TDT. In addition, we have begun our commercial launch of JOURNAVX for the treatment of acute pain, which received marketing approval in the U.S. in January 2025. We also continue to advance our pipeline of product candidates for the treatment of serious diseases outside of CF, SCD, TDT, and acute pain.
Our strategy is to combine transformative advances in the understanding of causal human biology and the science of therapeutics to discover and develop innovative medicines. This approach includes advancing multiple compounds or therapies from each program, spanning multiple modalities, into early clinical trials to obtain patient data that can inform selection of the most promising therapies for later-stage development, as well as to inform discovery and development efforts. We aim to rapidly follow our first-in-class therapies that achieve proof-of-concept with potential best-in-class candidates to provide durable clinical and commercial success.
In pursuit of new product candidates and therapies in specialty markets, we invest in research and development. We believe that pursuing research in diverse areas allows us to balance the risks inherent in product development and may provide product candidates that will form our pipeline in future years. To supplement our internal research programs, we acquire technologies and programs and collaborate with biopharmaceutical and technology companies, leading academic research institutions, government laboratories, foundations and other organizations, as needed, to advance research in our areas of therapeutic interest and to access technologies needed to execute on our strategy.
Discovery and development of a new pharmaceutical or biological product is a difficult and lengthy process that requires significant financial resources along with extensive technical and regulatory expertise. Across the industry, most potential drug or biological products never progress into development, and most products that advance into development never receive marketing approval. Our investments in product candidates are subject to considerable risks. We closely monitor our research and development activities, and frequently evaluate our pipeline programs in light of new data and scientific, business and commercial insights, with the objective of balancing risk and potential. This process can result in rapid changes in focus and priorities as new information becomes available and as we gain additional understanding of our ongoing programs and potential new programs, as well as those of our competitors. In addition, our product candidates must satisfy rigorous standards of safety and efficacy before they can be approved for sale by regulatory authorities. Our analysis of data obtained from nonclinical and clinical activities is subject to confirmation and interpretation by regulatory authorities, which could delay, limit or prevent regulatory approval.
Our business also requires ensuring appropriate manufacturing and supply of our products. As we advance our product candidates through clinical development toward commercialization and market and sell our approved products, we build and maintain our supply chain and quality assurance resources. We rely on a global network of third parties, including some in
China, and our internal capabilities to manufacture and distribute our products for commercial sale and post-approval clinical trials and to manufacture and distribute our product candidates for clinical trials. In addition to establishing supply chains for each newly approved product, we adapt our supply chain for existing products to include additional formulations or to increase scale of production for existing products as needed. Our foreign third-party manufacturers and suppliers may be subject to U.S. legislation, including the BIOSECURE Act, tariffs, sanctions, trade restrictions and other foreign regulatory requirements which could increase costs or reduce the supply of material available to us, or delay the procurement or supply of such material. The processes for biological and cell and genetic therapies can be more complex than those required for small molecule drugs and require additional investments in different systems, equipment, facilities and expertise. We are focused on ensuring the stability of the supply chains for our current products, as well as for our pipeline programs.
Sales of our products depend, to a large degree, on the extent to which our products are reimbursed by third-party payors, such as government health programs, commercial insurance and managed health care organizations. Reimbursement for our products, including our potential pipeline therapies, cannot be assured and may take significant periods of time to obtain. We dedicate substantial management and other resources to obtain and maintain appropriate levels of reimbursement for our products from third-party payors, including governmental organizations in the U.S. and ex-U.S. markets.
In the U.S., we have worked successfully with third-party payors to promptly obtain appropriate levels of reimbursement for our CF medicines. In addition, we are working with U.S. government and commercial payors with respect to CASGEVY and JOURNAVX. We anticipate broad access with government and commercial payors for CASGEVY in the U.S., and we have recently entered into multiple agreements with government and commercial health insurance providers to provide such access. For JOURNAVX in the U.S., we have been working with government and commercial payors pre- and post-approval to support rapid and broad access. We plan to continue to engage in discussions with numerous commercial insurers and managed health care organizations, along with government health programs that are typically managed by authorities in the individual states, to ensure that payors recognize the significant benefits that all our therapies provide and provide patients with appropriate levels of access to our medicines and therapies now and in the future. We cannot, however, predict how changes in the law, including through the Inflation Reduction Act of 2022 and passage of state laws (e.g., transparency laws and prescription drug affordability boards), will affect our ability to negotiate successfully with third-party payors and distribute our products. Similarly, in ex-U.S. markets, we seek government reimbursement for our medicines on a country-by-country or region-by-region basis, as required. This is necessary for each new medicine, as well as for label expansions for our current medicines. We are working with ex-U.S. payors with respect to CASGEVY, and we are pursuing long-term reimbursement agreements. We have secured reimbursed access for people with SCD or TDT in Saudi Arabia, Bahrain, Luxembourg, and England. In addition, the Italian Medicines Agency has approved early access for CASGEVY, on a case-by-case basis, to treat people with SCD and TDT. We expect to continue to focus significant resources to expand and maintain reimbursement for our CF medicines, CASGEVY, JOURNAVX, and, ultimately, our pipeline therapies, in U.S. and ex-U.S. markets.
Strategic Transactions
Acquisitions
As part of our business strategy, we seek to acquire technologies, products, product candidates and other businesses that are aligned with our corporate and research and development strategies and complement and advance our ongoing research and development efforts. We have acquired multiple biotechnology companies over the last several years and expect to continue to identify and evaluate such opportunities. The accounting for these acquisitions can vary significantly based on whether we conclude the transactions represent business combinations or asset acquisitions. In May 2024, we acquired Alpine Immune Sciences, Inc. (“Alpine”) for approximately $5.0 billion in cash. Alpine’s lead molecule, povetacicept, has shown potential to treat multiple diseases or conditions and become a pipeline-in-a-product. We accounted for the Alpine transaction as an asset acquisition because povetacicept represented substantially all of the fair value of the gross assets that we acquired. As a result, $4.4 billion of the fair value attributed to povetacicept was expensed as AIPR&D in the second quarter of 2024. In 2019 and 2022, we acquired Semma Therapeutics, Inc. (“Semma”) and ViaCyte, Inc. (“ViaCyte”), respectively, pursuant to which we established and accelerated the development of our T1D program. We accounted for each of these acquisitions as a business combination.
Collaboration and In-Licensing Arrangements
We enter into arrangements with third parties, including collaboration and licensing arrangements, for the development, manufacture and commercialization of products, product candidates and other technologies that have the potential to complement our ongoing research and development efforts.
Over the last several years, we entered into collaboration agreements with a number of companies, including CRISPR Therapeutics AG (“CRISPR”), Entrada Therapeutics, Inc. (“Entrada”), and Moderna, Inc.
Generally, when we in-license a technology or product candidate, we make upfront payments to the collaborator, assume the costs of the program and/or agree to make contingent payments, which could consist of milestone, royalty and option payments. Most of these collaboration payments are expensed as AIPR&D, including a $75.0 million milestone due to Entrada in the first quarter of 2024. These payments were expensed to AIPR&D because they were primarily attributable to acquired in-process research and development for which there was no alternative future use. However, depending on many factors, including the structure of the collaboration, the stage of development of the acquired technology, the significance of the in-licensed product candidate to the collaborator’s operations and the other activities in which our collaborators are engaged, the accounting for these transactions can vary significantly. We expect to continue to identify and evaluate collaboration and licensing opportunities that may be similar to or different from the collaborations and licenses that we have engaged in previously.
Acquired In-Process Research and Development Expenses
In the first quarter of 2025 and 2024, our AIPR&D included $19.8 million and $76.8 million, respectively, related to upfront, contingent milestone, or other payments pursuant to our business development transactions, including the asset acquisitions, collaborations, and licenses of third-party technologies described above. Please refer to Note B, “Collaboration, License and Other Arrangements,” for further information regarding our asset acquisitions, collaborations and in-license agreements.
Out-licensing Arrangements
We also have out-licensed certain development programs to collaborators who are leading the development or commercialization of these programs, either globally or within certain geographic regions.
In January 2025, we entered into a collaboration agreement with Zai Lab Limited (“Zai”) for the development and commercialization of povetacicept in mainland China, Hong Kong SAR, Macau SAR, Taiwan region and Singapore. Zai is responsible for the povetacicept clinical trials and regulatory submissions in the licensed territories. Zai will also be responsible for commercialization activities in the licensed territories, if povetacicept becomes an approved product. We are eligible to receive certain regulatory milestone payments and tiered royalties on future net sales of povetacicept in the region of focus for Zai.
RESULTS OF OPERATIONS
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2025 | 2024 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages and per share amounts) | |||||||||||||||||||||||||||||||||||
| Total revenues | $ | 2,770.2 | $ | 2,690.6 | 3% | ||||||||||||||||||||||||||||||
| Acquired in-process research and development expenses | 19.8 | 76.8 | (74)% | ||||||||||||||||||||||||||||||||
| Intangible asset impairment charge | 379.0 | — | N/A | ||||||||||||||||||||||||||||||||
| Other operating costs and expenses | 1,741.3 | 1,474.3 | 18% | ||||||||||||||||||||||||||||||||
| Income from operations | 630.1 | 1,139.5 | (45)% | ||||||||||||||||||||||||||||||||
| Other non-operating income, net | 100.3 | 139.6 | (28)% | ||||||||||||||||||||||||||||||||
| Provision for income taxes | 84.1 | 179.5 | (53)% | ||||||||||||||||||||||||||||||||
| Net income | $ | 646.3 | $ | 1,099.6 | (41)% | ||||||||||||||||||||||||||||||
| Net income per diluted common share | $ | 2.49 | $ | 4.21 | |||||||||||||||||||||||||||||||
| Diluted shares used in per share calculations | 259.5 | 261.1 | |||||||||||||||||||||||||||||||||
Total Revenues
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2025 | 2024 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| TRIKAFTA/KAFTRIO | $ | 2,535.5 | $ | 2,483.6 | 2% | ||||||||||||||||||||||||||||||
| ALYFTREK | 53.9 | — | N/A | ||||||||||||||||||||||||||||||||
| Other product revenues | 170.8 | 207.0 | (17)% | ||||||||||||||||||||||||||||||||
| Product revenues, net | 2,760.2 | 2,690.6 | 3% | ||||||||||||||||||||||||||||||||
| Other revenues | 10.0 | — | N/A | ||||||||||||||||||||||||||||||||
| Total revenues | $ | 2,770.2 | $ | 2,690.6 | 3% | ||||||||||||||||||||||||||||||
Product Revenues, Net
In the first quarter of 2025, our net product revenues increased by $69.6 million, or 3%, as compared to the first quarter of 2024, primarily due to the continued performance of TRIKAFTA/KAFTRIO and initial contributions from ALYFTREK. In the first quarter of 2025, "Other product revenues" included $14.2 million from CASGEVY, and an insignificant amount from JOURNAVX. In the first quarter of 2024, there were no revenues for these products.
Our net product revenues from the U.S. and from ex-U.S. markets were as follows:
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2025 | 2024 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| United States | $ | 1,653.5 | $ | 1,519.9 | 9% | ||||||||||||||||||||||||||||||
| ex-U.S. | 1,106.7 | 1,170.7 | (5)% | ||||||||||||||||||||||||||||||||
| Product revenues, net | $ | 2,760.2 | $ | 2,690.6 | 3% |
Our net product revenues increased 9% in the U.S. due to higher net realized pricing and continued strong patient demand. Our net product revenues decreased 5% outside the U.S. primarily due to an expected decline in product revenues in Russia, where we are continuing to experience a violation of our intellectual property rights.
Other Revenues
In the first quarter of 2025, “Other revenues” included a $10.0 million upfront payment we received from our collaboration agreement with Zai.
Operating Costs and Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2025 | 2024 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Cost of sales | $ | 363.0 | $ | 342.6 | 6% | ||||||||||||||||||||||||||||||
| Research and development expenses | 979.7 | 789.1 | 24% | ||||||||||||||||||||||||||||||||
| Acquired in-process research and development expenses | 19.8 | 76.8 | (74)% | ||||||||||||||||||||||||||||||||
| Selling, general and administrative expenses | 396.4 | 342.7 | 16% | ||||||||||||||||||||||||||||||||
| Intangible asset impairment charge | 379.0 | — | N/A | ||||||||||||||||||||||||||||||||
| Change in fair value of contingent consideration | 2.2 | (0.1) | ** | ||||||||||||||||||||||||||||||||
| Total costs and expenses | $ | 2,140.1 | $ | 1,551.1 | 38% | ||||||||||||||||||||||||||||||
| ** Not meaningful |
Cost of Sales
Our cost of sales primarily consists of third-party royalties payable on net sales of our CF products as well as the cost of producing inventories. Pursuant to our agreement with the Cystic Fibrosis Foundation, our tiered third-party royalties on sales of ALYFTREK, TRIKAFTA/KAFTRIO, SYMDEKO/SYMKEVI, KALYDECO, and ORKAMBI, calculated as a percentage of net sales, range from the single digits to the sub-teens, with lower royalties on sales of ALYFTREK and TRIKAFTA/KAFTRIO than for our other products.
In the first quarter of 2025, our cost of sales increased $20.4 million, or 6%, as compared to the first quarter of 2024, primarily due to increased sales volume and changes in product mix. Our cost of sales as a percentage of our net product revenues was 13% in each of the first quarter of 2025 and 2024.
Research and Development Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2025 | 2024 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Research expenses | $ | 206.1 | $ | 196.1 | 5% | ||||||||||||||||||||||||||||||
| Development expenses | 773.6 | 593.0 | 30% | ||||||||||||||||||||||||||||||||
| Total research and development expenses | $ | 979.7 | $ | 789.1 | 24% |
Our research and development expenses include internal and external costs incurred for research and development of our products and product candidates. We do not assign our internal costs, such as salary and benefits, stock-based compensation expense, laboratory supplies and other direct expenses and infrastructure costs, to individual products or product candidates, because the employees within our research and development groups typically are deployed across multiple research and development programs. We assign external costs of services provided to us by clinical research organizations and other outsourced research by individual program. Our internal costs are greater than our external costs. All research and development costs for our products and product candidates are expensed as incurred.
Since January 2023, we have incurred approximately $7.8 billion in research and development expenses associated with product discovery and development. The successful development of our product candidates is highly uncertain and subject to a number of risks. In addition, the duration of clinical trials may vary substantially according to the type, complexity and novelty of the product candidate and the disease indication being targeted. The FDA and comparable agencies in foreign countries impose substantial requirements on the introduction of therapeutic pharmaceutical products, typically requiring lengthy and detailed laboratory and clinical testing procedures, sampling activities and other costly and time-consuming procedures. Data obtained from nonclinical and clinical activities at any step in the testing process may be adverse and lead to discontinuation or redirection of development activities. Data obtained from these activities also are susceptible to varying interpretations, which could delay, limit or prevent regulatory approval. The duration and cost of discovery, nonclinical studies and clinical trials may vary significantly over the life of a project and are difficult to predict. Therefore, accurate and meaningful estimates of the ultimate costs to bring our product candidates to market are not available.
Any estimates regarding development and regulatory timelines for our product candidates are highly subjective and subject to change. Until we have data from Phase 3 clinical trials, we cannot make a meaningful estimate regarding when, or if, a clinical development program will generate revenues and cash flows.
Research Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2025 | 2024 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Research Expenses: | |||||||||||||||||||||||||||||||||||
| Salary and benefits | $ | 53.1 | $ | 53.0 | 0% | ||||||||||||||||||||||||||||||
| Stock-based compensation expense | 22.3 | 29.9 | (25)% | ||||||||||||||||||||||||||||||||
| Outsourced services and other direct expenses | 73.1 | 64.4 | 14% | ||||||||||||||||||||||||||||||||
| Infrastructure costs | 57.6 | 48.8 | 18% | ||||||||||||||||||||||||||||||||
| Total research expenses | $ | 206.1 | $ | 196.1 | 5% | ||||||||||||||||||||||||||||||
Our research expenses have been increasing over the last several years as we invested in our pipeline and expanded our cell and genetic therapy capabilities, which resulted in increased outsourced services and other direct expenses and infrastructure costs in the first quarter of 2025 as compared to the first quarter of 2024. We expect to continue to invest in our research programs with a focus on creating transformative medicines for serious diseases.
Development Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2025 | 2024 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Development Expenses: | |||||||||||||||||||||||||||||||||||
| Salary and benefits | $ | 195.9 | $ | 170.1 | 15% | ||||||||||||||||||||||||||||||
| Stock-based compensation expense | 77.8 | 89.5 | (13)% | ||||||||||||||||||||||||||||||||
| Outsourced services and other direct expenses | 379.7 | 236.1 | 61% | ||||||||||||||||||||||||||||||||
| Infrastructure costs | 120.2 | 97.3 | 24% | ||||||||||||||||||||||||||||||||
| Total development expenses | $ | 773.6 | $ | 593.0 | 30% | ||||||||||||||||||||||||||||||
Our development expenses increased by $180.6 million, or 30%, in the first quarter of 2025 as compared to the first quarter of 2024, due to continued investments in internal headcount and infrastructure to support multiple mid- and late-stage clinical development programs, driving increases in our outsourced services and other direct expenses. Our outsourced services and other direct expenses increased as compared to the first quarter of 2024 primarily due to the acquisition of our povetacicept program in the second quarter of 2024, expansion of our T1D study of VX-880, and next-generation pain product candidates.
Our stock-based compensation expenses, including those recorded as research and development expenses, have historically fluctuated and is expected to continue to fluctuate from one period to another primarily due to changes in the probability of achieving milestones associated with our performance-based awards.
Acquired In-process Research and Development Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2025 | 2024 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Acquired in-process research and development expenses | $ | 19.8 | $ | 76.8 | (74)% | ||||||||||||||||||||||||||||||
AIPR&D in the first quarter of 2025 was primarily related to milestone payments. AIPR&D in the first quarter of 2024 was primarily related to the $75.0 million milestone to Entrada. Our AIPR&D has historically fluctuated, and is expected to continue to fluctuate, from one period to another due to upfront, contingent milestone, and other payments pursuant to our existing and future business development transactions, including collaborations, licenses of third-party technologies, and asset acquisitions.
Selling, General and Administrative Expenses
| Three Months Ended March 31, | |||||||||||||||||||||||||||||||||||
| 2025 | 2024 | Change | |||||||||||||||||||||||||||||||||
| (in millions, except percentages) | |||||||||||||||||||||||||||||||||||
| Selling, general and administrative expenses | $ | 396.4 | $ | 342.7 | 16% |
Selling, general and administrative expenses increased by 16% in the first quarter of 2025 as compared to the first quarter of 2024 primarily due to increased commercial investment to support the launch of JOURNAVX.
Intangible Asset Impairment Charge
In the first quarter of 2025, based on results from a Phase 1/2 clinical trial evaluating our VX-264 clinical program in patients with T1D, we concluded that VX-264 will not be advancing further in clinical development. Based on this event, we
performed an interim impairment test on the fair value of our VX-264 indefinite-lived in-process research and development asset that we acquired from Semma Therapeutics, Inc. in 2019. As a result, we recorded a full intangible asset impairment charge of $379.0 million associated with VX-264.
Contingent Consideration
The fair value of our contingent consideration increased by $2.2 million in the first quarter of 2025 and decreased by $0.1 million in the first quarter of 2024.
Other Non-Operating Income (Expense), Net
Interest Income
Interest income decreased from $181.2 million in the first quarter of 2024 to $120.9 million in the first quarter of 2025, primarily due to decreased cash equivalents and available-for-sale debt securities following our acquisition of Alpine in the second quarter of 2024 and decreased market interest rates compared to the first quarter of 2024. Our future interest income is dependent on the amount of, and prevailing market interest rates on, our outstanding cash equivalents and available-for-sale debt securities.
Interest Expense
Interest expense was $3.0 million and $10.4 million in the first quarter of 2025 and 2024, respectively. The majority of our interest expense in these periods was related to imputed interest expense associated with our finance leases, including interest expense imputed on our corporate headquarters leases in Boston in the first quarter of 2024. Our corporate headquarters leases were amended in the third quarter of 2024 resulting in a classification change from finance to operating leases, which are recorded entirely within operating expenses in our condensed consolidated statements of operations.
Other Income (Expense), Net
Other income (expense), net were expenses of $17.6 million and $31.2 million in the first quarter of 2025 and 2024, respectively. These amounts primarily related to net unrealized losses resulting from changes in the fair value of certain of our strategic equity investments, which consist of investments in our collaborators that may be public or privately-held companies. To the extent that we continue to hold strategic equity investments in publicly traded biotechnology companies, we expect that our other income (expense), net will continue to fluctuate in future periods due to the volatility in the stock prices of these companies that impacts the fair value of our investments. As of March 31, 2025, the fair value of our investments in publicly traded companies was $21.6 million.
Income Taxes
Our effective tax rate fluctuates from period to period due to the global nature of our operations. The factors that most significantly impact our effective tax rate include changes in tax laws, variability in the amount and allocation of our taxable earnings among multiple jurisdictions, the amount and characterization of our research and development expenses, the levels of certain deductions and credits, adjustments to the value of our uncertain tax positions, acquisitions and third-party collaboration and licensing transactions.
We recorded provisions for income taxes of $84.1 million and $179.5 million in the first quarter of 2025 and 2024, respectively. Our effective tax rate of 11.5% in the first quarter of 2025 was lower than the U.S. statutory rate primarily due to excess tax benefits related to stock-based compensation. Our effective tax rate of 14.0% in the first quarter of 2024 was lower than the U.S. statutory rate primarily due to changes in our unrecognized tax positions as well as excess tax benefits related to stock-based compensation.
LIQUIDITY AND CAPITAL RESOURCES
The following table summarizes the components of our financial condition as of March 31, 2025 and December 31, 2024:
| As of March 31, 2025 | As of December 31, 2024 | Change | |||||||||||||||
| (in millions, except percentages) | |||||||||||||||||
| Cash, cash equivalents and marketable securities: | |||||||||||||||||
| Cash and cash equivalents | $ | 4,674.7 | $ | 4,569.6 | |||||||||||||
| Marketable securities | 1,526.5 | 1,546.3 | |||||||||||||||
| Long-term marketable securities | 5,156.5 | 5,107.9 | |||||||||||||||
| Total cash, cash equivalents and marketable securities | $ | 11,357.7 | $ | 11,223.8 | 1% | ||||||||||||
| Working Capital: | |||||||||||||||||
| Total current assets | $ | 10,008.8 | $ | 9,596.4 | 4% | ||||||||||||
| Total current liabilities | (3,783.2) | (3,564.6) | 6% | ||||||||||||||
| Total working capital | $ | 6,225.6 | $ | 6,031.8 | 3% |
Working Capital
As of March 31, 2025, total working capital was $6.2 billion, which represented an increase of $193.8 million, or 3%, compared to December 31, 2024, primarily due to increased cash and cash equivalents resulting from our operations and increased inventories following the regulatory approvals of ALYFTREK and JOURNAVX.
Cash Flows
| Three Months Ended March 31, | |||||||||||
| 2025 | 2024 | ||||||||||
| (in millions) | |||||||||||
| Net cash provided by (used in): | |||||||||||
| Operating activities | $ | 818.9 | $ | 1,306.6 | |||||||
| Investing activities | $ | (55.8) | $ | (2,136.4) | |||||||
| Financing activities | $ | (680.4) | $ | (357.5) |
Operating Activities
Cash provided by operating activities decreased from $1.3 billion in the first quarter of 2024 to $818.9 million in the first quarter of 2025, primarily due to the timing of tax payments and accruals related to our product revenue contracts.
Investing Activities
Cash used in investing activities was $55.8 million in the first quarter of 2025, primarily related to purchases of property and equipment. Cash used in investing activities was $2.1 billion in the first quarter of 2024. The largest portion of our investing activities in the first quarter of 2024 were net purchases of available-for-sale debt securities.
Financing Activities
Cash used in financing activities were $680.4 million and $357.5 million in the first quarter of 2025 and 2024, respectively. Our financing activities in each of these periods were primarily related to repurchases of our common stock pursuant to our share repurchase program and payments related to our employee stock benefit plans.
Sources and Uses of Liquidity
We intend to rely on our existing cash, cash equivalents and current marketable securities together with our operating profitability as our primary source of liquidity. We expect that cash flows from our product sales together with our cash, cash equivalents and current marketable securities will be sufficient to fund our operations for at least the next twelve months. The
adequacy of our available funds to meet our future operating and capital requirements will depend on many factors, including our future sales of currently marketed products, and the potential introduction of one or more new product candidates to the market, our business development activities, and the number, breadth and cost of our research and development programs.
Credit Facilities & Financing Strategy
We may borrow up to a total of $500.0 million pursuant to a revolving credit facility that we entered into in July 2022 and could repay and reborrow amounts under this revolving credit agreement without penalty. Subject to certain conditions, we could request that the borrowing capacity be increased by an additional $500.0 million, for a total of $1.0 billion. Negative covenants in our credit agreement could prohibit or limit our ability to access this source of liquidity. As of March 31, 2025, the facility was undrawn, and we were in compliance with these covenants.
We may also raise additional capital by borrowing under credit agreements, through public offerings or private placements of our securities, or securing new collaborative agreements or other methods of financing. We will continue to manage our capital structure and will consider all financing opportunities, whenever they may occur, that could strengthen our long-term liquidity profile. There can be no assurance that any such financing opportunities will be available on acceptable terms, if at all.
Future Capital Requirements
We have significant future capital requirements, including:
-
Expected operating expenses to conduct research and development activities, manufacture and commercialize our existing and future products, and to operate our organization.
-
Cash that we pay for income taxes.
-
Royalties we pay related to sales of our CF products.
-
Facility, operating and finance lease obligations.
-
Firm purchase obligations related to our supply and manufacturing processes.
In addition, other potential significant future capital requirements may include:
-
We have entered into certain agreements with third parties that include the funding of certain research, development, manufacturing and commercialization efforts. Certain of our transactions, including collaborations, licensing arrangements, and asset acquisitions, include the potential for future milestone and royalty payments by us upon the achievement of pre-established developmental and regulatory targets and/or commercial targets. Other transactions include the potential for future lease-related expenses and other costs. Our obligation to fund these research and development and commercialization efforts and to pay these potential milestones, expenses and royalties is contingent upon continued involvement in the programs and/or the lack of any adverse events that could cause their discontinuance. We may enter into additional agreements, including acquisitions, collaborations, licensing arrangements and equity investments, which require additional capital.
-
To the extent we borrow amounts under our existing credit agreement, we would be required to repay any outstanding principal amounts in 2027.
-
As of March 31, 2025, we had $964.4 million remaining authorization available under our $3.0 billion Share Repurchase Program that our Board of Directors approved in February 2023. This program does not have an expiration date and can be discontinued at any time. We expect to fund these programs through a combination of cash on hand and cash generated by operations.
There have not been any material changes to our future capital requirements disclosed in our Annual Report on Form 10-K for the year ended December 31, 2024, which was filed with the Securities and Exchange Commission, or SEC, on February 13, 2025.
CRITICAL ACCOUNTING POLICIES AND ESTIMATES
Our discussion and analysis of our financial condition and results of operations are based upon our condensed consolidated financial statements prepared in accordance with generally accepted accounting principles in the U.S. The
preparation of these financial statements requires us to make certain estimates and assumptions that affect the reported amounts of assets and liabilities, the disclosure of contingent assets and liabilities at the date of the condensed consolidated financial statements and the reported amounts of revenues and expenses during the reported periods. These items are monitored and analyzed by management for changes in facts and circumstances, and material changes in these estimates could occur in the future. Changes in estimates are reflected in reported results for the period in which the change occurs. We base our estimates on historical experience and various other assumptions that we believe to be reasonable under the circumstances. Actual results may differ from our estimates if past experience or other assumptions do not turn out to be substantially accurate. During the three months ended March 31, 2025, there were no material changes to our critical accounting policies as reported in our Annual Report on Form 10-K for the year ended December 31, 2024, which was filed with the SEC on February 13, 2025.
RECENT ACCOUNTING PRONOUNCEMENTS
For a discussion of recent accounting pronouncements, please refer to Note A, “Basis of Presentation and Accounting Policies.”
Previous: Item 1. Financial Statements · Next: Item 3. Quantitative and Qualitative Disclosures About Market Risk