 filings.tools
Biogen 10-K 2014-12-31
Filed 2015-02-04. 21 sections, 597K characters. Original on sec.gov · Markdown · JSON
Cover and table of contents
10-K 1 biib-20141231x10k.htm 10-K
UNITED STATES SECURITIES AND EXCHANGE COMMISSION
Washington, D.C. 20549
Form 10-K
| þ | ANNUAL REPORT PURSUANT TO SECTION 13 OR 15(d) OF THE SECURITIES EXCHANGE ACT OF 1934 |
| For the fiscal year ended December 31, 2014 |
or
| ¨ | TRANSITION REPORT PURSUANT TO SECTION 13 OR 15(d) OF THE SECURITIES EXCHANGE ACT OF 1934 |
Commission file number: 0-19311
BIOGEN IDEC INC.
(Exact name of registrant as specified in its charter)
| Delaware | 33-0112644 | |
| (State or other jurisdiction of incorporation or organization) | (I.R.S. Employer Identification No.) |
225 Binney Street, Cambridge, Massachusetts 02142
(617) 679-2000
(Address, including zip code, and telephone number, including area code, of Registrant’s principal executive offices)
Securities registered pursuant to Section 12(b) of the Act:
| Title of Each Class | Name of Each Exchange on Which Registered | |
| Common Stock, $0.0005 par value | The Nasdaq Global Select Market |
Securities registered pursuant to Section 12(g) of the Act:
None
Indicate by check mark if the registrant is a well-known seasoned issuer, as defined in Rule 405 of the Securities Act. Yes þ No ¨
Indicate by check mark if the registrant is not required to file reports pursuant to Section 13 or 15(d) of the Act. Yes ¨ No þ
Indicate by check mark whether the registrant (1) has filed all reports required to be filed by Section 13 or 15(d) of the Securities Exchange Act of 1934 during the preceding 12 months (or for such shorter period that the registrant was required to file such reports), and (2) has been subject to such filing requirements for the past 90 days. Yes þ No ¨
Indicate by check mark whether the registrant has submitted electronically and posted on its corporate Web site, if any, every Interactive Data File required to be submitted and posted pursuant to Rule 405 of Regulation S-T during the preceding 12 months (or for such shorter period that the registrant was required to submit and post such files): Yes þ No ¨
Indicate by check mark if disclosure of delinquent filers pursuant to Item 405 of Regulation S-K is not contained herein, and will not be contained, to the best of the registrant’s knowledge, in definitive proxy or information statements incorporated by reference in Part III of this Form 10-K or any amendment to this Form 10-K. þ
Indicate by check mark whether the registrant is a large accelerated filer, an accelerated filer, a non-accelerated filer, or a smaller reporting company. See the definitions of “large accelerated filer,” “accelerated filer” and “smaller reporting company” in Rule 12b-2 of the Exchange Act.
| Large accelerated filer þ | Accelerated filer ¨ | Non-accelerated filer ¨ | Smaller reporting company ¨ | |||
| (Do not check if a smaller reporting company) |
Indicate by check mark whether the registrant is a shell company (as defined in Rule 12b-2 of the Act). Yes ¨ No þ
The aggregate market value of the registrant’s common stock held by non-affiliates of the registrant (without admitting that any person whose shares are not included in such calculation is an affiliate) computed by reference to the price at which the common stock was last sold as of the last business day of the registrant’s most recently completed second fiscal quarter was $74,386,280,444.
As of January 30, 2015, the registrant had 234,614,474 shares of common stock, $0.0005 par value, outstanding.
DOCUMENTS INCORPORATED BY REFERENCE
Portions of the definitive proxy statement for our 2015 Annual Meeting of Stockholders are incorporated by reference into Part III of this report.
BIOGEN IDEC INC.
ANNUAL REPORT ON FORM 10-K
For the Year Ended December 31, 2014
TABLE OF CONTENTS
NOTE REGARDING FORWARD-LOOKING STATEMENTS
This report contains forward-looking statements that are being made pursuant to the provisions of the Private Securities Litigation Reform Act of 1995 (the “Act”) with the intention of obtaining the benefits of the “Safe Harbor” provisions of the Act. These forward-looking statements may be accompanied by such words as “anticipate,” “believe,” “could,” “estimate,” “expect,” “forecast,” “intend,” “may,” “plan,” “potential,” “project,” “target,” “will” and other words and terms of similar meaning. Reference is made in particular to forward-looking statements regarding:
| • | the anticipated amount, timing and accounting of revenues, contingent payments, milestone, royalty and other payments under licensing, collaboration or acquisition agreements, tax positions and contingencies, collectability of receivables, pre-approval inventory, cost of sales, research and development costs, compensation and other expenses, amortization of intangible assets, foreign currency exchange risk and impairment assessments; |
| • | the potential impact of increased product competition in the multiple sclerosis (MS), hemophilia and oncology markets; |
| • | the timing, outcome and impact of administrative, regulatory, legal and other proceedings related to patents and other proprietary and intellectual property rights, tax audits, assessments and settlements, pricing matters, sales and promotional practices, product liability and other matters; |
| • | patent terms, patent term extensions, patent office actions, and expected availability and period of regulatory exclusivity; |
| • | the costs, timing, potential approval and therapeutic scope of the development and commercialization of our pipeline products and the expected timing of related filings and regulatory actions; |
| • | our intent to commit resources for research and development opportunities; |
| • | our manufacturing capacity and use of third party contract manufacturing organizations to provide manufacturing services; |
| • | the drivers for growing our business, including our plans to pursue business development and research opportunities; |
| • | the potential impact of healthcare reform in the U.S., implementation of provisions of the Patient Protection and Affordable Care Act (also known as the Affordable Care Act or PPACA), and measures being taken worldwide designed to reduce healthcare costs to constrain the overall level of government expenditures, including the impact of pricing actions in Europe and elsewhere, and reduced reimbursement for our products; |
| • | lease commitments, purchase obligations and the timing and satisfaction of other contractual obligations; |
| • | the impact of the continued uncertainty and deterioration of the credit and economic conditions in certain countries in Europe and our collection of accounts receivable in such countries; |
| • | our ability to finance our operations and business initiatives and obtain funding for such activities; and |
| • | the impact of new laws and accounting standards. |
These forward-looking statements involve risks and uncertainties, including those that are described in the “Risk Factors” section of this report and elsewhere within this report that could cause actual results to differ materially from those reflected in such statements. You should not place undue reliance on these statements. Forward-looking statements speak only as of the date of this report. We do not undertake any obligation to publicly update any forward-looking statements.
NOTE REGARDING COMPANY AND PRODUCT REFERENCES
Throughout this report, “Biogen Idec,” the “Company,” “we,” “us” and “our” refer to Biogen Idec Inc. and its consolidated subsidiaries. References to “RITUXAN” refer to both RITUXAN (the trade name for rituximab in the U.S., Canada and Japan) and MabThera (the trade name for rituximab outside the U.S., Canada and Japan), and “ANGIOMAX” refers to both ANGIOMAX (the trade name for bivalirudin in the U.S., Canada and Latin America) and ANGIOX (the trade name for bivalirudin in Europe).
NOTE REGARDING TRADEMARKS
ALPROLIX®, AVONEX®, ELOCTATE®, RITUXAN®, TECFIDERA®, and TYSABRI® are registered trademarks of Biogen Idec. ELOCTATM, FUMADERMTM, PLEGRIDYTM and ZINBRYTATM are trademarks of Biogen Idec. The following are trademarks of the respective companies listed: ACTEMRA® - Chugai Seiyaku Kabushiki Kaisha; ADVATE® - Baxter International Inc.; ANGIOMAX® and ANGIOXTM - The Medicines Company; ARZERRA® - Glaxo Group Limited; BENLYSTA® - GlaxoSmithKline Intellectual Property Limited; AUBAGIO® - Sanofi Societe Anonyme France; BENEFIX® - Genetics Institute LLC; BETASERON®- Bayer Pharma AG; CIMZIA® - UCB Pharma, S.A.; COPAXONE® - Teva Pharmaceutical Industries Limited; ENBREL® - Immunex Corporation; EXTAVIA® and GILENYA® - Novartis AG; FAMPYRATM - Acorda Therapeutics, Inc.; GAZYVA® - Genentech, Inc.; HELIXATE® - CSL Behring LLC; HUMIRA® - AbbVie Biotechnology Ltd.; IMBRUVICA® - Pharmacyclics, Inc.; KOGENATE® - Bayer AG; LEMTRADA® - Genzyme Corporation; ORENCIA® - Bristol-Myers Squibb Company; REBIF® - Ares Trading S.A.; REMICADE® - Janssen Biotech, Inc.; RIXUBIS - Baxter International Inc.; SIMPONI® and SIMPONI ARIATM - Johnson & Johnson; TREANDA® - Cephalon, Inc.; XELJANZ® - Pfizer Inc.; XYNTHA® - Wyeth LLC; and ZYDELIG® - Gilead Sciences.
PART I
Item 1. Business
Overview
Biogen Idec is a global biopharmaceutical company focused on discovering, developing, manufacturing and delivering therapies for neurological, autoimmune and hematologic disorders. Our principal marketed products include AVONEX, PLEGRIDY, TECFIDERA, TYSABRI, and FAMPYRA for multiple sclerosis (MS), ALPROLIX for hemophilia B and ELOCTATE for hemophilia A. We also collaborate on the development and commercialization of RITUXAN for the treatment of non-Hodgkin's lymphoma, chronic lymphocytic leukemia and other conditions and share profits and losses for GAZYVA which is approved for the treatment of chronic lymphocytic leukemia.
We are focused on discovering and developing new therapies that improve the lives of patients having diseases with high unmet medical needs. We support our mission through the commitment of significant resources to research and development programs and business development opportunities, particularly within areas of our scientific, manufacturing and technical expertise - neurology, immunology and hematology, and scientific adjacencies.
We were formed as a corporation in the State of California in 1985 under the name IDEC Pharmaceuticals Corporation and reincorporated as a Delaware corporation in 1997. In 2003, we acquired Biogen, Inc. and changed our corporate name to Biogen Idec Inc.
Marketed Products
The following charts show our product sales and unconsolidated joint business revenues by principal product and geography as a percentage of revenue for the years ended December 31, 2014, 2013 and 2012.
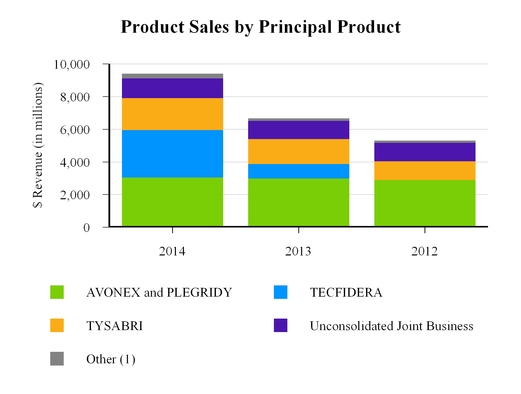
(1) Other includes FAMPYRA, ALPROLIX, ELOCTATE and FUMADERM.
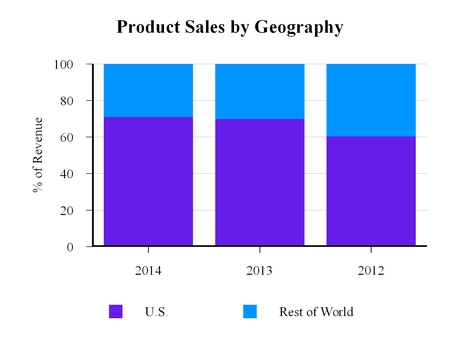
Product sales for AVONEX, TECFIDERA and TYSABRI and unconsolidated joint business revenues for RITUXAN each accounted for more than 10% of our total revenue for the years ended December 31, 2014 and 2013, and AVONEX, TYSABRI and RITUXAN each accounted for more than 10% of our total revenue for the year ended December 31, 2012. For additional financial information about our product and other revenues and geographic areas in which we operate, please read Note 25, Segment Information to our consolidated financial statements, Item 6. Selected Financial Data and Item 7. Management's Discussion and Analysis of Financial Condition and Results of Operations included in this report. A discussion of the risks attendant to our operations is set forth in the “Risk Factors” section of this report.
Multiple Sclerosis (MS) Products
We develop, manufacture and market a number of products designed to treat patients with MS. MS is a progressive neurological disease in which the body loses the ability to transmit messages along nerve cells, leading to a loss of muscle control, paralysis and, in some cases, death. Patients with active relapsing MS experience an uneven pattern of disease progression characterized by periods of stability that are interrupted by flare-ups of the disease after which the patient returns to a new baseline of functioning. Our MS products include:
| • | AVONEX (interferon beta-1a), an intramuscular injectable therapy, indicated for the treatment of patients with relapsing forms of MS. AVONEX is a recombinant form of the interferon beta protein produced in the body in response to viral infection. The principal markets for AVONEX are the U.S., United Kingdom, France, Germany, Italy and Spain. |
| • | PLEGRIDY (peginterferon beta-1a), a subcutaneous injectable therapy, indicated in the U.S. for the treatment of patients with relapsing forms of MS and in the European Union (E.U.) for relapsing-remitting MS (RRMS). PLEGRIDY received approval from the European Commission (EC) in July 2014 and the U.S. Food and Drug Administration (FDA) in August 2014. |
| • | TECFIDERA (dimethyl fumarate), an oral therapy indicated in the U.S. for the treatment of patients with relapsing forms of MS and in the EU for people with RRMS. TECFIDERA was approved by the FDA in March 2013 and the EC in February 2014. |
| • | TYSABRI (natalizumab), a monoclonal antibody approved in numerous countries as a monotherapy for the treatment of patients with relapsing forms of MS. TYSABRI is also approved in the U.S. to treat Crohn's disease, an inflammatory disease of the intestines. The principal markets for TYSABRI in MS are the U.S., the United Kingdom, France, Germany, Italy and Spain. TYSABRI was approved in Japan in March 2014. |
| • | FAMPYRA (prolonged-release fampridine tablets), is indicated for the improvement of walking ability in adult patients with MS. FAMPYRA is a prolonged-release tablet formulation of the drug fampridine. We have a license from Acorda Therapeutics, Inc. (Acorda) to develop and commercialize FAMPYRA in all markets outside the U.S. Our principal markets for FAMPRYA are France, Germany, Spain and Canada. For information about our agreement with Acorda, please read Note 20, Collaborative and Other Relationships to our consolidated financial statements included in this report. |
Hemophilia Products
We develop, manufacture and market products designed to treat patients with hemophilia A and B. Hemophilia A is caused by having substantially reduced or no Factor VIII activity and hemophilia B is caused by having substantially reduced or no Factor IX activity, each of which is needed for normal blood clotting. People with hemophilia A and B experience bleeding episodes that may cause pain, irreversible joint damage and life-threatening hemorrhages. Prophylactic infusions of Factor VIII or Factor IX, as applicable, temporarily replace clotting factor necessary to control bleeding and prevent new bleeding episodes. Our products for hemophilia A and B include:
| • | ALPROLIX [Coagulation Factor IX (Recombinant), Fc Fusion Protein], a recombinant DNA-derived, coagulation Factor IX concentrate indicated in the U.S. for treatment in adults and children with hemophilia B for control and prevention of bleeding episodes, perioperative management and routine prophylaxis to prevent or reduce the frequency of bleeding episodes. ALPROLIX was approved by the FDA in March 2014 and in Japan in June 2014. |
| • | ELOCTATE [Antihemophilic Factor (Recombinant), Fc Fusion Protein], a recombinant DNA-derived, antihemophilic factor indicated in the U.S. for treatment in adults and children with hemophilia A for control and prevention of bleeding episodes, perioperative management and routine prophylaxis to prevent or reduce the frequency of bleeding episodes. ELOCTATE was approved by the FDA in June 2014 and in Japan in December 2014. |
We collaborate with Swedish Orphan Biovitrum AB (Sobi) to jointly develop and commercialize Factor VIII and Factor IX hemophilia products, including ELOCTATE and ALPROLIX. For information about our agreement with Sobi, please read Note 20, Collaborative and Other Relationships to our consolidated financial statements included in this report.
Genentech Collaboration
We collaborate with Genentech, Inc. (Genentech), a wholly-owned member of the Roche Group, on the development and commercialization of RITUXAN. We also share operating profits and losses relating to GAZYVA with Genentech in the U.S. The Roche Group and its sub-licensees maintain sole responsibility for the development, manufacturing and commercialization of GAZYVA in the U.S. For information about our unconsolidated joint business and agreement with Genentech, please read Note 1, Summary of Significant Accounting Policies and Note 20, Collaborative and Other Relationships to our consolidated financial statements included in this report.
| • | RITUXAN (rituximab), a widely prescribed monoclonal antibody use |
Showing the first 8K of 94K characters. Open the full section
Item 1A. Risk Factors
We are substantially dependent on revenues from our principal products.
Our current revenues depend upon continued sales of our principal products, TECFIDERA, AVONEX, TYSABRI, and RITUXAN. We may be substantially dependent on sales from our principal products for many years, including an increasing reliance on sales and growth of TECFIDERA as we continue to expand into additional markets. Any negative developments relating to any of these products, including the following, and as discussed in greater detail in these “Risk Factors”, may adversely affect our revenues and results of operations or could cause a decline in our stock price:
| • | safety or efficacy issues; |
| • | the introduction or greater acceptance of competing products; |
| • | constraints and additional pressures on product pricing or price increases, due to a number of factors, including governmental or regulatory requirements, increased competition, or changes in reimbursement policies and practices of payors and other third parties; or |
| • | adverse legal, administrative, regulatory or legislative developments. |
If we fail to compete effectively, our business and market position would suffer.
The biopharmaceutical industry and the markets in which we operate are intensely competitive. We compete in the marketing and sale of our products, the development of new products and processes, the acquisition of rights to new products with commercial potential and the hiring and retention of personnel. We compete with biotechnology and pharmaceutical companies that have a greater number of products on the market and in the product pipeline, greater financial and other resources and other technological or competitive advantages. One or more of our competitors may benefit from significantly greater sales and marketing capabilities, may develop products that are accepted more widely than ours or may receive patent protection that dominates, blocks or adversely affects our product development or business.
Our products are also susceptible to competition from generics and biosimilars in many markets. Generic versions of drugs and biosimilars are likely to be sold at substantially lower prices than branded products. Accordingly, the introduction of generic or biosimilar versions of our marketed products likely would significantly reduce both the price that we receive for such marketed products and the volume of products that we sell, which may have an adverse impact on our results of operations.
In the MS market, we face intense competition as the number of products and competitors continues to expand. Due to our significant reliance on sales of our MS products, our business may be harmed if we are unable to successfully compete in the MS market. More specifically, our ability to compete and maintain and grow our share in the MS market may be adversely affected due to a number of factors, including:
| • | the introduction of more efficacious, safer, less expensive or more convenient alternatives to our MS products, including our own products; |
| • | the introduction of lower-cost biosimilars, follow-on products or generic versions of branded MS products sold by our competitors, and the possibility of future competition from generic versions or related prodrug derivatives or from off-label use by physicians of therapies indicated for other conditions to treat MS patients; |
| • | patient dynamics, including the size of the patient population and our ability to attract new patients to our therapies; |
| • | damage to physician and patient confidence in any of our MS products or to our sales and reputation as a result of label changes or adverse experiences or events that may occur with patients treated with our MS products; |
| • | inability to obtain appropriate pricing and reimbursement for our MS products compared to our competitors in key international markets; or |
| • | our ability to obtain and maintain patent, data or market exclusivity for our MS products. |
Similarly, the hemophilia treatment market is highly competitive, with current treatments marketed by companies that have substantially greater financial resources and marketing expertise. Our ability to successfully compete in the hemophilia market and gain share in this market may be adversely affected due to a number of reasons, including:
| • | difficulty in penetrating this market if our therapies are not regarded as offering substantial benefits over current treatments; |
| • | the introduction by other companies of longer-lasting or more efficacious, safer, less expensive or more convenient treatments than our therapies; |
| • | our limited marketing experience within the hemophilia treatment market, which may impact our ability to develop well-established relationships with the associated medical and scientific community; |
| • | our failure to receive positive pediatric data from our ongoing global pediatric studies, which is required for filing our planned MAA for ALPROLIX with the EMA; or |
| • | if one of several companies that are working to develop additional treatments for hemophilia obtains marketing approval of its treatment in the E.U. before we do, our application with the EMA could be barred under operation of the EMA’s Orphan Medicines Regulation. |
If we are unable to obtain and maintain adequate protection for our data, intellectual property and other proprietary rights, our business may be harmed.
Our success depends in part on our ability to obtain and defend patent and other intellectual property rights that are important to the commercialization of our products and product candidates. The degree of patent protection that will be afforded to our products and processes in the U.S. and in other important markets remains uncertain and is dependent upon the scope of protection decided upon by the patent offices, courts and lawmakers in these countries. We can provide no assurance that we will successfully obtain or preserve patent protection for the technologies incorporated into our products and processes, or that the protection obtained will be of sufficient breadth and degree to protect our commercial interests in all countries where we conduct business. If we cannot prevent others from exploiting our inventions, we will not derive the benefit from them that we currently expect. Furthermore, we can provide no assurance that our products will not infringe patents or other intellectual property rights held by third parties.
We also rely on regulatory exclusivity for protection of our products. Implementation and enforcement of regulatory exclusivity, which may consist of regulatory data protection and market protection, varies widely from country to country. Failure to qualify for regulatory exclusivity, or failure to obtain or maintain the extent or duration of such protections that we expect in each of the markets for our products, could affect our revenue for our products or our decision on whether to market our products in a particular country or countries or could otherwise have an adverse impact on our results of operations.
Litigation, interference, oppositions or other proceedings have been and may in the future be necessary in some instances to determine the validity and scope of certain of our proprietary rights, and in other instances to determine the validity, scope or non-infringement of certain patent rights claimed by third parties to be pertinent to the manufacture, use or sale of our products. We may also face challenges to our patent and regulatory protections covering our products by manufacturers of generics and biosimilars that may choose to launch or attempt to launch their products before the expiration of our patent or regulatory exclusivity. Litigation, interference, oppositions or other similar types of proceedings are unpredictable and may be protracted, expensive and distracting to management. The outcome of such proceedings could adversely affect the validity and scope of our patent or other proprietary rights, hinder our ability to manufacture and market our products, require us to seek a license for the infringed product or technology or result in the assessment of significant monetary damages against us that may exceed amounts, if any, accrued in our financial statements. An adverse determination in a judicial or administrative proceeding or a failure to obtain necessary licenses could prevent us from manufacturing or selling our products. Furthermore, payments under any licenses that we are able to obtain would reduce our profits derived from the covered products and services.
Sales of our products depend, to a significant extent, on adequate coverage, pricing and reimbursement from third party payors, which are subject to increasing and intense pressure from political, social, competitive and other sources. Our inability to maintain adequate coverage, or a reduction in pricing or reimbursement, could have an adverse effect on our business, revenues and results of operations, and could cause a decline in our stock price.
Sales of our products are dependent, in large part, on the availability and extent of coverage, pricing and reimbursement from government health administration authorities, private health insurers and other organizations, and drug prices are under significant scrutiny in the markets where our products are prescribed. Our ability to set the price for our products can vary significantly from country to country and as a result so can the price of our products, and we may continue to face increasing pressure to lower the prices for our products in many markets. Changes in government regulations or private third-party payors' reimbursement policies, as well as pressure by employers on private health insurance plans to reduce costs, may reduce pricing and reimbursement for our products and adversely affect our future results. In addition, when a new medical product is approved, the availability of government and private reimbursement for that product is uncertain, as is the pricing and amount for which that product will be reimbursed. We also cannot predict the availability, pricing or amount of reimbursement for our product candidates. Our failure to maintain adequate coverage, pricing, or reimbursement for our products would have an adverse effect on our business, revenues and results of operation, could curtail or eliminate our ability to adequately fund research and development programs for the discovery and commercialization of new products, and could cause a decline in our stock price.
In the U.S., federal and state legislatures, health agencies and third-party payors continue to focus on containing the cost of health care. Legislative and regulatory proposals and enactments to reform health care insurance programs could significantly influence the manner in which our products are prescribed and purchased. For example, provisions of the PPACA have resulted in changes in the way health care is paid for by both governmental and private insurers, including increased rebates owed by manufacturers under the Medicaid Drug Rebate Program, annual fees and taxes on manufacturers of certain branded prescription drugs, the requirement that manufacturers participate in a discount program for certain outpatient drugs under Medicare Part D and the expansion of the number of hospitals eligible for discounts under Section 340B of the Public Health Service Act. These changes have had and are expected to continue to have a significant impact on our business.
Managed care organizations continue to seek price discounts and, in some cases, to impose restrictions on the coverage of particular drugs. For example, health insurers, pharmacy benefit managers and other payors may seek price discounts or rebates in connection with the placement of our products on their formularies. They could also impose restrictions on access to our products, and could even choose to exclude coverage of our products entirely.
There is also significant economic pressure on state budgets that may result in states increasingly seeking to achieve budget savings through mechanisms that limit coverage or payment for our drugs. In recent years, some states have considered legislation that would control the prices of drugs, including laws to allow importation of pharmaceutical products from lower cost jurisdictions outside the U.S. State Medicaid programs are increasingly requesting manufacturers to pay supplemental rebates and requiring prior authorization by the state program for use of any drug for which supplemental rebates are not being paid. Government efforts to reduce Medicaid expenses may lead to increased use of managed care organizations by Medicaid programs. This may result in managed care organizations influencing prescription decisions for a larger segment of the population and a corresponding constraint on prices and reimbursement for our products. In addition, under the PPACA, as states implement their health care marketplaces or operate under the federal exchange, the impact on drug manufacturers, including us, will depend in part on the formulary and benefit design decisions made by insurance sponsors or plans participating in these programs. It is possible that we may need to provide discounts or rebates to such plans in order to maintain favorable formulary access for our products for this patient population, which could have an adverse impact on our sales and results of operations.
In the European Union and some other international markets, the government provides health care at low cost to consumers and regulates pharmaceutical prices, patient eligibility or reimbursement levels to control costs for the government-sponsored health care system. Many countries have announced or implemented measures to reduce health care costs to constrain their overall level of government expenditures. These measures vary by country and may include, among other things, patient access restrictions, suspensions on price increases, prospective and possibly retroactive price reductions and other recoupments and increased mandatory discounts or rebates, recoveries of past price increases, and greater importation of drugs from lower-cost countries to higher-cost countries. These measures have negatively impacted our revenues, and may continue to adversely affect our revenues and results of operations in the future. In addition, certain countries set prices by reference to the prices in other countries where our products are marketed. Thus, our inability to secure adequate prices in a particular country may not only limit the marketing of our products within that country, but may also adversely affect our ability to obtain acceptable prices in other markets. This may create the opportunity for third party cross-border trade or influence our decision to sell or not to sell a product, thus adversely affecting our geographic expansion plans and revenues.
Adverse safety events or restrictions on use and safety warnings for our products can negatively affect our business, product sales and stock price.
Adverse safety events involving our marketed products may have a negative impact on our business. Discovery of safety issues with our products could create product liability and could cause additional regulatory scrutiny and requirements for additional labeling, withdrawal of products from the market, and the imposition of fines or criminal penalties. Adverse safety events may also damage physician and patient confidence in our products and our reputation. Any of these could result in liabilities, loss of revenue, material write-offs of inventory, material impairments of intangible assets, goodwill and fixed assets, material restructuring charges and other adverse impacts on our results of operations.
Regulatory authorities are making greater amounts of stand-alone safety information directly available to the public through periodic safety update reports, patient registries and other reporting requirements. The reporting of adverse safety events involving our products or products similar to ours and public rumors about such events may increase claims against us and may also cause our product sales or stock price to decline or experience periods of volatility.
Restrictions on use or significant safety warnings that may be required to be included in the label of our products, such as the risk of developing progressive multifocal leukoencephalopathy (PML), a serious brain infection, in the label for TYSABRI and in the U.S. label for TECFIDERA, may significantly reduce expected revenues for those products and require significant expense and management time.
Our long-term success depends upon the successful development, license or acquisition of new products and additional indications for existing products.
Our long-term viability and growth will depend upon the successful development of new products and technologies from our research and development activities, including those licensed or acquired from third parties and biosimilars developed through our joint venture with Samsung Biologics, and approval of additional indications for our existing products. The sustainability of growth in our business is dependent in part upon our ability to continue to build a strong early and mid-stage pipeline of product candidates in our core competencies as well as additional areas of unmet need. Product development is very expensive and involves a high degree of risk. Only a small number of research and development programs result in the commercialization of a product. Success in preclinical work or early stage clinical trials does not ensure that later stage or larger scale clinical trials will be successful or that regulatory approval will be obtained. Further, it is possible that we may experience delays, uncertainties or difficulties developing biosimilars as the legislative and regulatory pathways for approval of biosimilars continue to evolve in many markets.
Conducting clinical trials is a complex, time-consuming and expensive process. Our ability to complete our clinical trials in a timely fashion depends in large part on a number of key factors including protocol design, regulatory and institutional review board approval, patient enrollment rates, and compliance with extensive current Good Clinical Practices. We have opened clinical sites and are enrolling patients in a number of countries where our experience is more limited. In most cases, we use the services of third party clinical trial providers and third party contract research organizations, or CROs, to carry out most of our clinical trial related activities and accurately report their results, which may impact our ability to control the timing, conduct, expense and quality of our clinical trials. One CRO has responsibility for substantially all of our clinical trial related activities and reporting. If our CROs do not successfully carry out their activities or meet expected deadlines, our trials may be delayed. We may also need to replace our CROs. Although we believe that there are a number of other third-party CROs we could engage to continue these activities, the replacement of an existing CRO may result in delay of the affected trials or otherwise adversely affect our efforts to obtain regulatory approvals and commercialize our drug candidates.
If we fail to adequately manage the design, execution and regulatory aspects of our large, complex and diverse clinical trials, our studies and any potential regulatory approvals may be delayed, or we may fail to gain approvals for our product candidates. Clinical trials may indicate that our product candidates lack efficacy, have harmful side effects or raise safety or other concerns that may significantly reduce the likelihood of regulatory approval, result in significant restrictions on use and safety warnings in the approved label, adversely affect placement within the treatment paradigm, or otherwise significantly diminish the commercial potential of the product candidate. Also, positive results in a registrational trial may not be replicated in any subsequent confirmatory trials. Even if later stage clinical trials are successful, regulatory authorities may disagree with our view of the data or require additional studies, may disagree with trial design or the endpoints employed in the trials, may fail to approve the facilities or the processes used to manufacture a product candidate, may fail to approve or delay approval of our product candidates, dosing or delivery methods, companion devices or may otherwise grant marketing approval that is more restricted than anticipated, including indications covering narrow patient populations and the imposition of safety monitoring or educational requirements or risk evaluation and mitigation strategies. The occurrence of any such events could result in the incurrence of significant costs and expenses, have an adverse effect on our business, including our financial condition and results of operations, or cause our stock price to decline or experience periods of volatility.
Even if we are able to successfully develop new products or indications, we may make a strategic decision to discontinue development of such product or indication if, for example, we believe commercialization will be difficult relative to other opportunities in our pipeline.
We depend on relationships with collaborators and other third-parties for product and royalty revenue, and the development, commercialization and marketing of certain products, which are outside of our full control.
We rely on a number of significant collaborative relationships for product and royalty revenue, and the development, commercialization, and marketing of certain of our products and product candidates. Reliance on collaborative relationships subjects us to a number of risks, including:
| • | we may be unable to control the resources our collaborator devotes to our programs or products; |
| • | disputes may arise with respect to ownership of rights to technology developed with our collaboration partner, and the underlying contract with our collaborator may fail to provide significant protection or may fail to be effectively enforced if the collaborator fails to perform; |
| • | our collaborators’ interests may not always be aligned with our interests and they may not market a product in the same manner or to the same extent that we would, which could adversely affect our revenues; |
| • | collaborations often require the parties to cooperate, and failure to do so effectively could adversely affect product sales by our collaborators or the clinical development or regulatory approvals of products under joint control or could result in termination of the research, development or commercialization of product candidates or result in litigation or arbitration; and |
| • | any failure on the part of our collaborators to comply with applicable laws and regulatory requirements in the marketing, sale and maintenance of the market authorization of our products or to fulfill any responsibilities they may have to protect and enforce any intellectual property rights underlying our products could have an adverse effect on our revenues as well as involve us in possible legal proceedings. |
Given these risks, there is considerable uncertainty regarding the success of our current and future collaborative efforts. If these efforts fail, our product development or commercialization of new products could be delayed or revenues from products could decline.
Manufacturing issues could substantially increase our costs, limit supply of our products and reduce our revenues.
The process of manufacturing our products is complex, highly regulated and subject to numerous risks, including:
| • | The manufacturing process for our products is extremely susceptible to product loss due to contamination, oxidation, equipment failure or improper installation or operation of equipment, or vendor or operator error. Even minor deviations from normal manufacturing processes could result in reduced production yields, product defects and other supply disruptions. If microbial, viral or other contaminations are discovered in our products or manufacturing facilities, we may need to close our manufacturing facilities for an extended period of time to investigate and remediate the contaminant. |
| • | We rely on third party suppliers and manufacturers for, among other things: manufacturing of RITUXAN and GAZYVA; the majority of our clinical and commercial requirements for TECFIDERA and other small molecule products and product candidates; raw materials and supplies for production of products we manufacture; delivery devices such as syringes and auto-injectors; drug product and fill-finish operations; the majority of our final product storage; and a substantial portion of our packaging operations. In addition, due to the unique manner in which our products are manufactured, we rely on single source providers of several raw materials and manufacturing supplies. These third parties are independent entities subject to their own unique operational and financial risks that are outside of our control. These third parties may not perform their obligations in a timely and cost-effective manner or in compliance with applicable regulations, and they may be unable or unwilling to increase production capacity commensurate with demand for our existing or future products. Finding alternative providers could take a significant amount of time and involve significant expense due to the specialized nature of the services and the need to obtain regulatory approval of any significant changes to our suppliers or manufacturing methods. We cannot be certain that we could reach agreement with alternative providers or that the FDA or other regulatory authorities would approve our use of such alternatives. |
| • | We rely on our manufacturing facilities in Cambridge, Massachusetts, RTP, North Carolina and Hillerød, Denmark for the production of drug substance for certain of our large molecule products and product candidates, including AVONEX, TYSABRI, PLEGRIDY, ZINBRYTA, ALPROLIX and ELOCTATE. Our global bulk supply of these products and product candidates depends on the uninterrupted and efficient operation of these facilities, which could be adversely affected by equipment failures, labor shortages, natural disasters, power failures and numerous other factors. |
| • | We and our third party providers are generally required to maintain compliance with current Good Manufacturing Practices and other stringent requirements and are subject to inspections by the FDA and comparable agencies in other jurisdictions to confirm such compliance. Any delay, interruption or other issues that arise in the manufacture, fill-finish, packaging, or storage of our products as a result of a failure of our facilities or the facilities or operations of third parties to pass any regulatory agency inspection could significantly impair our ability to develop and commercialize our products. Significant noncompliance could also result in the imposition of monetary penalties or other civil or criminal sanctions and damage our reputation. |
Any adverse developments affecting our manufacturing operations or the operations of our third-party suppliers and manufacturers may result in shipment delays, inventory shortages, lot failures, product withdrawals or recalls, or other interruptions in the commercial supply of our products. We may also have to take inventory write-offs and incur other charges and expenses for products that fail to meet specifications, undertake costly remediation efforts or seek more costly manufacturing alternatives. Such developments could increase our manufacturing costs, cause us to lose revenue or market share as patients and physicians turn to competing therapeutics, diminish our profitability or damage our reputation.
While we believe we currently have sufficient manufacturing capacity to meet our near-term manufacturing requirements, it is probable that we would need additional manufacturing capacity to support future clinical and commercial manufacturing requirements for product candidates in our pipeline, if such candidates are successful and approved. Due to the long lead times necessary for the expansion of manufacturing capacity, it is possible that we may incur significant costs to build or acquire additional facilities or obtain third party contract manufacturers in advance of product demand and sales. If we are unable to adequately and timely manufacture and supply our products and product candidates, our business may be harmed.
Our business may be adversely affected if we do not manage our current growth and do not successfully execute our growth initiatives.
We have experienced significant growth in our headcount and operations, which has placed, and will continue to place, significant demands on our management and our operational and financial infrastructure. We anticipate further growth through both internal development projects as well as external opportunities, which may include the acquisition, partnering and in-licensing of products, technologies and companies or the entry into strategic alliances and collaborations. The availability of high quality development opportunities is limited and competitive, and we are not certain that we will be able to identify candidates that we and our shareholders consider suitable or complete transactions on terms that are acceptable to us and our shareholders. In order to pursue such opportunities, we may require significant additional financing, which may not be available to us on favorable terms, if at all. Even if we are able to successfully identify and complete acquisitions and other strategic alliances and collaborations, we may face unanticipated costs or liabilities in connection with the transaction or we may not be able to integrate them or take full advantage of them or otherwise realize the benefits that we expect.
To manage our current and future potential growth effectively, we need to continue to enhance our operational, financial and management processes and to expand, train and manage our employee base. Our growth is also dependent upon our ability to attract and retain qualified scientific, information technology, manufacturing, sales and marketing and executive personnel and to develop and maintain relationships with qualified clinical researchers and key distributors in a highly competitive environment. Supporting our growth initiatives and the further development of our existing products and potential new products in our pipeline will require significant capital expenditures and management resources, including investments in research and development, sales and marketing, manufacturing capabilities and other areas of our business. If we do not successfully manage our current growth and do not successfully execute our growth initiatives, then our business and financial results may be adversely affected and we may incur asset impairment or restructuring charges.
If we fail to comply with the extensive legal and regulatory requirements affecting the health care industry, we could face increased costs, penalties and a loss of business.
Our activities, and the activities of our collaborators, distributors and other third party providers, are subject to extensive government regulation and oversight both in the U.S. and in foreign jurisdictions. The FDA and comparable agencies in other jurisdictions directly regulate many of our most critical business activities, including the conduct of preclinical and clinical studies, product manufacturing, advertising and promotion, product distribution, adverse event reporting and product risk management. Our interactions in the U.S. or abroad with physicians and other health care providers that prescribe or purchase our products are also subject to government regulation designed to prevent fraud and abuse in the sale and use of the products and place greater restrictions on the marketing practices of health care companies. Health care companies such as ours are facing heightened scrutiny of their relationships with health care providers from anti-corruption enforcement officials. In addition, we along with many other pharmaceutical and biotechnology companies have been the target of lawsuits and investigations alleging violations of government regulation, including claims asserting submission of incorrect pricing information, impermissible off-label promotion of pharmaceutical products, payments intended to influence the referral of health care business, submission of false claims for government reimbursement, antitrust violations, or violations related to
environmental matters. These risks may be heightened as we continue to expand our global operations and introduce additional products to the market.
Regulations governing the health care industry are subject to change, with possibly retroactive effect, including:
| • | new laws, regulations or judicial decisions, or new interpretations of existing laws, regulations or decisions, related to health care availability, pricing or marketing practices, compliance with wage and hour laws and other employment practices, method of delivery, payment for health care products and services, compliance with health information and data privacy and security laws and regulations, tracking and reporting payments and other transfers of value made to physicians and teaching hospitals, extensive anti-bribery and anti-corruption prohibitions, product serialization and labeling requirements, and used product take-back requirements; |
| • | changes in the FDA and foreign regulatory approval processes that may delay or prevent the approval of new products and result in lost market opportunity; |
| • | requirements that provide for increased transparency of clinical trial results and quality data, such as the EMA’s recently enacted clinical transparency policy, which could impact our ability to protect trade secrets and competitively-sensitive information contained in approval applications or could be misinterpreted leading to reputational damage, misperception or legal action which could harm our business; and |
| • | changes in FDA and foreign regulations that may require additional safety monitoring, labeling changes, restrictions on product distribution or use, or other measures after the introduction of our products to market, which could increase our costs of doing business, adversely affect the future permitted uses of approved products, or otherwise adversely affect the market for our products. |
Examples of previously enacted and possible future changes in laws that could adversely affect our business include the enactment in the U.S. of health care reform, potential regulations easing the entry of competing biosimilars in the marketplace, new legislation or implementation of existing statutory provisions on importation of lower-cost competing drugs from other jurisdictions, enhanced penalties for and investigations into non-compliance with U.S. fraud and abuse laws, and compliance with the Physician Payment Sunshine Act in the U.S. and similar foreign rules and regulations that require collection and reporting of payments or other transfers of value made to physicians and teaching hospitals.
Violations of governmental regulation may be punishable by criminal and civil sanctions against us, including fines and civil monetary penalties and exclusion from participation in government programs, including Medicare and Medicaid, as well as against executives overseeing our business. In addition to penalties for violation of laws and regulations, we could be required to repay amounts we received from government payors, or pay additional rebates and interest if we are found to have miscalculated the pricing information we have submitted to the government. Whether or not we have complied with the law, an investigation into alleged unlawful conduct could increase our expenses, damage our reputation, divert management time and attention and adversely affect our business.
A breakdown or breach of our information technology systems could subject us to liability or interrupt the operation of our business.
We are increasingly dependent upon information technology systems and data. Our computer systems continue to increase in multitude and complexity due to the growth in our business, making them potentially vulnerable to breakdown, malicious intrusion and random attack. Likewise, data privacy or security breaches by individuals authorized to access our information technology systems or others may pose a risk that sensitive data, including intellectual property, trade secrets or personal information belonging to us, our patients, customers or other business partners, may be exposed to unauthorized persons or to the public. Cyber-attacks are increasing in their frequency, sophistication and intensity. While we continue to build and improve our information systems and infrastructure and believe we have taken appropriate security measures to minimize these risks to our data and information technology systems, there can be no assurance that our efforts will prevent breakdowns or breaches in our systems that could adversely affect our business.
Our sales and operations are subject to the risks of doing business internationally.
We are increasing our presence in international markets, particularly emerging markets, subjecting us to many risks that could adversely affect our business and revenues, such as:
| • | the inability to obtain necessary foreign regulatory or pricing approvals of products in a timely manner; |
| • | collectability of accounts receivable; |
| • | fluctuations in foreign currency exchange rates, in particular a strengthening of the U.S. dollar versus foreign currencies; |
| • | difficulties in staffing and managing international operations; |
| • | the imposition of governmental controls; |
| • | less favorable intellectual property or other applicable laws; |
| • | increasingly complex standards for complying with foreign laws and regulations that may differ substantially from country to country and may conflict with corresponding U.S. laws and regulations; |
| • | the far-reaching anti-bribery and anti-corruption legislation in the U.K., including the U.K. Bribery Act 2010, and elsewhere and escalation of investigations and prosecutions pursuant to such laws; |
| • | compliance with complex import and export control laws; |
| • | restrictions on direct investments by foreign entities and trade restrictions; |
| • | greater political or economic instability; and |
| • | changes in tax laws and tariffs. |
In addition, our international operations are subject to regulation under U.S. law. For example, the Foreign Corrupt Practices Act prohibits U.S. companies and their representatives from offering, promising, authorizing or making payments to foreign officials for the purpose of obtaining or retaining business abroad. In many countries, the health care professionals we regularly interact with may meet the definition of a foreign government official for purposes of the Foreign Corrupt Practices Act. Failure to comply with domestic or foreign laws could result in various adverse consequences, including: possible delay in approval or refusal to approve a product; recalls, seizures or withdrawal of an approved product from the market; disruption in the supply or availability of our products or suspension of export or import privileges; the imposition of civil or criminal sanctions; the prosecution of executives overseeing our international operations; and damage to our reputation. Any significant impairment of our ability to sell products outside of the U.S. could adversely impact our business and financial results.
Our effective tax rate may fluctuate and we may incur obligations in tax jurisdictions in excess of accrued amounts.
As a global biopharmaceutical company, we are subject to taxation in numerous countries, states and other jurisdictions. As a result, our effective tax rate is derived from a combination of applicable tax rates in the various places that we operate. In preparing our financial statements, we estimate the amount of tax that will become payable in each of such places. Our effective tax rate, however, may be different than experienced in the past due to numerous factors, including changes in the mix of our profitability from country to country, the results of examinations and audits of our tax filings, adjustments to the value of our uncertain tax positions, changes in accounting for income taxes and changes in tax laws. Any of these factors could cause us to experience an effective tax rate significantly different from previous periods or our current expectations.
In addition, our inability to secure or sustain acceptable arrangements with tax authorities and previously enacted or future changes in the tax laws, among other things, may result in tax obligations in excess of amounts accrued in our financial statements.
In the U.S., there are several proposals under consideration to reform tax law, including proposals that may reduce or eliminate the deferral of U.S. income tax on our unrepatriated earnings, penalize certain transfer pricing structures, and reduce or eliminate certain foreign or domestic tax credits or deductions. Our future reported financial results may be adversely affected by tax law changes which restrict or eliminate certain foreign tax credits or our ability to deduct expenses attributable to foreign earnings, or otherwise affect the treatment of our unrepatriated earnings.
In addition to U.S. tax reform proposals, the adoption of some or all of the recommendations set forth in the Organization for Economic Co-operation and Development’s project on “Base Erosion and Profit Shifting” (BEPS) by tax authorities in the countries in which we operate, could negatively impact our effective tax rate. These recommendations focus on payments from affiliates in high tax jurisdictions to affiliates in lower tax jurisdictions and the activities that give rise to a taxable presence in a particular country.
Our operating results are subject to significant fluctuations.
Our quarterly revenues, expenses and net income (loss) have fluctuated in the past and are likely to fluctuate significantly in the future due to the risks described in these “Risk Factors” as well as the timing of charges and expenses that we may take. We have recorded, or may be required to record, charges that include:
| • | the cost of restructurings; |
| • | impairments with respect to investments, fixed assets and long-lived assets, including in-process R&D and other intangible assets; |
| • | inventory write-downs for failed quality specifications, charges for excess or obsolete inventory and charges for inventory write downs relating to product suspensions, expirations or recalls; |
| • | bad debt expenses and increased bad debt reserves; |
| • | outcomes of litigation and other legal or administrative proceedings, regulatory matters and tax matters; |
| • | milestone payments under license and collaboration agreements; and |
| • | payments in connection with acquisitions and other business development activities. |
Our revenues are also subject to foreign exchange rate fluctuations due to the global nature of our operations. We recognize foreign currency gains or losses arising from our operations in the period in which we incur those gains or losses. Although we have foreign currency forward contracts to hedge specific forecasted transactions denominated in foreign currencies, our efforts to reduce currency exchange losses may not be successful. As a result, currency fluctuations among our reporting currency, the U.S. dollar, and the currencies in which we do business will affect our operating results, often in unpredictable ways. Our net income may also fluctuate due to the impact of charges we may be required to take with respect to foreign currency hedge transactions. In particular, we may incur higher than expected charges from hedge ineffectiveness or from the termination of a hedge relationship.
Our operating results during any one period do not necessarily suggest the anticipated results of future periods.
Our investments in properties may not be fully realized.
We own or lease real estate primarily consisting of buildings that contain research laboratories, office space, and manufacturing operations. For strategic or other operational reasons, we may decide to further consolidate or co-locate certain aspects of our business operations or dispose of one or more of our properties, some of which may be located in markets that are experiencing high vacancy rates and decreasing property values. If we determine that the fair value of any of our owned properties is lower than their book value we may not realize the full investment in these properties and incur significant impairment charges. If we decide to fully or partially vacate a leased property, such as we did following our 2013 relocation of our corporate headquarters from Weston, Massachusetts to Cambridge, Massachusetts, we may incur significant cost, including lease termination fees, rent expense in excess of sublease income and impairment of leasehold improvements. In addition, in the event we expand our manufacturing capacity and we do not fully utilize our manufacturing facilities, this may result in idle time at facilities or substantial excess manufacturing capacity. Any of these events may have an adverse impact on our results of operations.
Our portfolio of marketable securities is subject to market, interest and credit risk that may reduce its value.
We maintain a portfolio of marketable securities for investment of our cash. Changes in the value of our portfolio of marketable securities could adversely affect our earnings. In particular, the value of our investments may decline due to increases in interest rates, downgrades of the bonds and other securities included in our portfolio, instability in the global financial markets that reduces the liquidity of securities included in our portfolio, declines in the value of collateral underlying the securities included in our portfolio, and other factors. Each of these events may cause us to record charges to reduce the carrying value of our investment portfolio or sell investments for less than our acquisition cost. Although we attempt to mitigate these risks through diversification of our investments and continuous monitoring of our portfolio's overall risk profile, the value of our investments may nevertheless decline.
We may not be able to access the capital and credit markets on terms that are favorable to us.
We may seek access to the capital markets to supplement our existing funds and cash generated from operations for working capital, capital expenditure and debt service requirements, and other business initiatives. The capital and credit markets have experienced extreme volatility and disruption which leads to uncertainty and liquidity issues for both borrowers and investors. In the event of adverse capital and credit market conditions, we may be unable to obtain capital market financing
on favorable terms. Changes in credit ratings issued by nationally recognized credit rating agencies could adversely affect our cost of financing and have an adverse effect on the market price of our securities.
Our business involves environmental risks, which include the cost of compliance and the risk of contamination or injury.
Our business and the business of several of our strategic partners involve the controlled use of hazardous materials, chemicals, biologics and radioactive compounds. Although we believe that our safety procedures for handling and disposing of such materials comply with state and federal standards, there will always be the risk of accidental contamination or injury. If we were to become liable for an accident, or if we were to suffer an extended facility shutdown, we could incur significant costs, damages and penalties that could harm our business. Manufacturing of our products and product candidates also requires permits from government agencies for water supply and wastewater discharge. If we do not obtain appropriate permits, or permits for sufficient quantities of water and wastewater, we could incur significant costs and limits on our manufacturing volumes that could harm our business.
The illegal distribution and sale by third parties of counterfeit versions of our products or stolen products could have a negative impact on our reputation and business.
Third parties might illegally distribute and sell counterfeit or unfit versions of our products, which do not meet our rigorous manufacturing, distribution and testing standards. A patient who receives a counterfeit or unfit drug may be at risk for a number of dangerous health consequences. Our reputation and business could suffer harm as a result of counterfeit or unfit drugs sold under our brand name. In addition, thefts of inventory at warehouses, plants or while in-transit, which are not properly stored and which are sold through unauthorized channels, could adversely impact patient safety, our reputation and our business.
The increasing use of social media platforms presents new risks and challenges.
Social media is increasingly being used to communicate about our products and the diseases our therapies are designed to treat. Social media practices in the biopharmaceutical industry continue to evolve and regulations relating to such use are not always clear. This evolution creates uncertainty and risk of noncompliance with regulations applicable to our business. For example, patients may use social media channels to comment on the effectiveness of a product or to report an alleged adverse event. When such disclosures occur, there is a risk that we fail to monitor and comply with applicable adverse event reporting obligations or we may not be able to defend the company or the public's legitimate interests in the face of the political and market pressures generated by social media due to restrictions on what we may say about our products. There is also a risk of inappropriate disclosure of sensitive information or negative or inaccurate posts or comments about us on any social networking website. If any of these events were to occur or we otherwise fail to comply with applicable regulations, we could incur liability, face overly restrictive regulatory actions or incur other harm to our business.
Some of our collaboration agreements contain change in control provisions that may discourage a third party from attempting to acquire us.
Some of our collaboration agreements include change in control provisions that could reduce the potential acquisition price an acquirer is willing to pay or discourage a takeover attempt that could be viewed as beneficial to shareholders. Upon a change in control, some of these provisions could trigger reduced milestone, profit or royalty payments to us or give our collaboration partner rights to terminate our collaboration agreement, acquire operational control or force the purchase or sale of the programs that are the subject of the collaboration.
Item 1B. Unresolved Staff Comments
None.
Item 2. Properties
Below is a summary of our owned and leased properties as of December 31, 2014.
Massachusetts
In Cambridge, Massachusetts, we own approximately 508,000 square feet of real estate space, consisting of a building that houses a research laboratory and a cogeneration plant totaling approximately 263,000 square feet and a building that contains research, development and quality laboratories which total approximately 245,000 square feet.
In addition, we lease a total of approximately 1,225,000 square feet in Massachusetts, which is summarized as follows:
| • | 822,000 square feet in Cambridge, Massachusetts, which is comprised of a 67,000 square foot biologics manufacturing facility and 755,000 square feet for our corporate headquarters, laboratory and additional office space; |
| • | 357,000 square feet of office space in Weston, Massachusetts, of which 175,000 square feet has been subleased through the remaining term of our lease agreement; and |
| • | 46,000 square feet of warehouse space in Somerville, Massachusetts. |
Our Massachusetts lease agreements expire at various dates through the year 2028.
North Carolina
In RTP, North Carolina, we own approximately 740,000 square feet of real estate space, which is summarized as follows:
| • | 357,000 square feet of laboratory and office space; |
| • | 175,000 square feet related to a large-scale biologics manufacturing facility; |
| • | 105,000 square feet related to a biologics manufacturing facility; |
| • | 60,000 square feet of warehouse space; and |
| • | 43,000 square feet related to a large-scale purification facility. |
In addition, we lease 48,000 square feet of a facility in RTP, North Carolina from Eisai to manufacture our and Eisai's oral solid dose products.
Denmark
We have a large-scale biologics manufacturing facility totaling approximately 228,000 square feet located in Hillerød, Denmark.
We also own approximately 306,000 square feet of additional space which is currently in use at this location and is summarized as follows:
| • | 139,000 square feet of warehouse, utilities and support space; |
| • | 70,000 square feet related to a label and packaging facility; |
| • | 47,000 square feet of administrative space; and |
| • | 50,000 square feet related to a laboratory facility. |
Other International
We lease office space in Zug, Switzerland, our international headquarters, the United Kingdom, Germany, France, Denmark, and numerous other countries. Our international lease agreements expire at various dates through the year 2023.
Item 3. Legal Proceedings
For a discussion of legal matters as of December 31, 2014, please read Note 21, Litigation to our consolidated financial statements included in this report, which is incorporated into this item by reference.
Item 4. Mine Safety Disclosures
Not applicable.
PART II
Item 5. Market for Registrant’s Common Equity, Related Stockholder Matters and Issuer Purchases of Equity Securities
Market and Stockholder Information
Our common stock trades on The NASDAQ Global Select Market under the symbol “BIIB.” The following table shows the high and low sales price for our common stock as reported by The NASDAQ Global Select Market for each quarter in the years ended December 31, 2014 and 2013:
| Common Stock Price | |||||||||||||||
| 2014 | 2013 | ||||||||||||||
| High | Low | High | Low | ||||||||||||
| First Quarter | $ | 358.89 | $ | 270.62 | $ | 192.92 | $ | 139.72 | |||||||
| Second Quarter | $ | 322.25 | $ | 272.02 | $ | 242.64 | $ | 191.80 | |||||||
| Third Quarter | $ | 349.00 | $ | 298.31 | $ | 248.95 | $ | 203.55 | |||||||
| Fourth Quarter | $ | 361.93 | $ | 290.85 | $ | 298.82 | $ | 221.07 |
As of January 30, 2015, there were approximately 816 stockholders of record of our common stock.
In addition, as of January 30, 2015, 30 stockholders of record of Biogen, Inc. common stock have yet to exchange their shares of Biogen, Inc. common stock for our common stock as contemplated by the merger of Biogen, Inc. and IDEC Pharmaceuticals Corporation in November 2003.
Dividends
We have not paid cash dividends since our inception. While we historically have not paid cash dividends and do not have a current intention to pay cash dividends, we continually review our capital allocation strategies, including, among other things, payment of cash dividends, stock repurchases, or acquisitions.
Issuer Purchases of Equity Securities
The following table summarizes our common stock repurchase activity during the fourth quarter of 2014:
| Period | Total Number of Shares Purchased (#) | Average Price Paid per Share ($) | Total Number of Shares Purchased as Part of Publicly Announced Programs (#) | Maximum Number of Shares That May Yet Be Purchased Under Our Programs | |||||||
| October 2014 | 7,862 | 297.72 | 7,771 | 2,939,869 | |||||||
| November 2014 | 1,395,731 | 314.38 | 1,395,731 | 1,544,138 | |||||||
| December 2014 | 279,982 | 305.98 | 279,982 | 1,264,156 | |||||||
| Total | 1,683,575 | 312.90 |
On February 11, 2011, we announced that our Board of Directors authorized the repurchase of up to 20.0 million shares of common stock. This authorization does not have an expiration date. As of December 31, 2014, approximately 18.7 million shares of our common stock at a cost of $2,770.0 million have been repurchased under this authorization and approximately 1.3 million shares of our common stock remain available for repurchase.
Stock Performance Graph
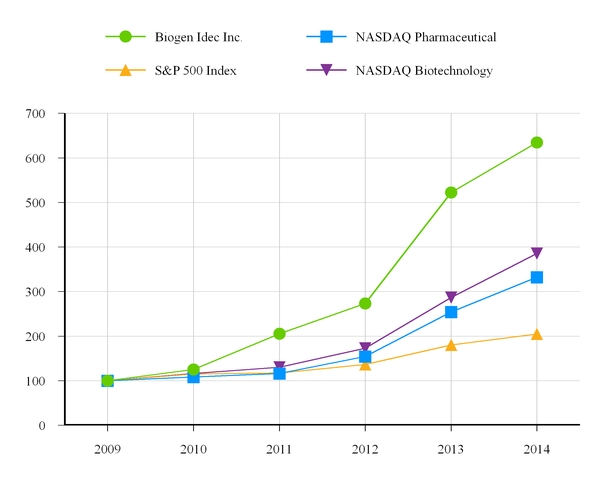
The graph below compares the five-year cumulative total stockholder return on our common stock, the S&P 500 Index, the Nasdaq Pharmaceutical Index and the Nasdaq Biotechnology Index assuming the investment of $100.00 on December 31, 2009 with dividends being reinvested. The stock price performance in the graph below is not necessarily indicative of future price performance.
| 2009 | 2010 | 2011 | 2012 | 2013 | 2014 | |||||||
| Biogen Idec Inc. | 100.00 | 125.33 | 205.70 | 273.59 | 522.56 | 634.49 | ||||||
| NASDAQ Pharmaceutical | 100.00 | 108.40 | 116.03 | 154.38 | 254.51 | 332.21 | ||||||
| S&P 500 Index | 100.00 | 115.06 | 117.49 | 136.30 | 180.44 | 205.14 | ||||||
| NASDAQ Biotechnology | 100.00 | 116.06 | 130.08 | 172.67 | 286.67 | 385.29 |
Item 6. Selected Financial Data
BIOGEN IDEC INC. AND SUBSIDIARIES
SELECTED FINANCIAL DATA
| For the Years Ended December 31, | |||||||||||||||||||
| 2014 | 2013 | 2012 | 2011 | 2010 | |||||||||||||||
| (In millions, except per share amounts) | (2) (4) | (2) (3) | (1) | ||||||||||||||||
| Results of Operations | |||||||||||||||||||
| Product revenues | $ | 8,203.4 | $ | 5,542.3 | $ | 4,166.1 | $ | 3,836.1 | $ | 3,470.1 | |||||||||
| Revenues from unconsolidated joint business | 1,195.4 | 1,126.0 | 1,137.9 | 996.6 | 1,077.2 | ||||||||||||||
| Other revenues | 304.5 | 263.9 | 212.5 | 215.9 | 169.1 | ||||||||||||||
| Total revenues | 9,703.3 | 6,932.2 | 5,516.5 | 5,048.6 | 4,716.4 | ||||||||||||||
| Total cost and expenses | 5,747.7 | 4,441.6 | 3,707.4 | 3,323.9 | 3,467.5 | ||||||||||||||
| Gain on sale of rights | 16.8 | 24.9 | 46.8 | — | — | ||||||||||||||
| Income from operations | 3,972.4 | 2,515.5 | 1,855.9 | 1,724.7 | 1,248.9 | ||||||||||||||
| Other income (expense), net | (25.8 | ) | (34.9 | ) | (0.7 | ) | (13.5 | ) | (19.0 | ) | |||||||||
| Income before income tax expense and equity in loss of investee, net of tax | 3,946.6 | 2,480.6 | 1,855.1 | 1,711.2 | 1,229.9 | ||||||||||||||
| Income tax expense | 989.9 | 601.0 | 470.6 | 444.5 | 331.3 | ||||||||||||||
| Equity in loss of investee, net of tax | 15.1 | 17.2 | 4.5 | — | — | ||||||||||||||
| Net income | 2,941.5 | 1,862.3 | 1,380.0 | 1,266.7 | 898.6 | ||||||||||||||
| Net income (loss) attributable to noncontrolling interests, net of tax | 6.8 | — | — | 32.3 | (106.7 | ) | |||||||||||||
| Net income attributable to Biogen Idec Inc. | $ | 2,934.8 | $ | 1,862.3 | $ | 1,380.0 | $ | 1,234.4 | $ | 1,005.2 | |||||||||
| Diluted Earnings Per Share | |||||||||||||||||||
| Diluted earnings per share attributable to Biogen Idec Inc. | $ | 12.37 | $ | 7.81 | $ | 5.76 | $ | 5.04 | $ | 3.94 | |||||||||
| Weighted-average shares used in calculating diluted earnings per share attributable to Biogen Idec Inc. | 237.2 | 238.3 | 239.7 | 245.0 | 254.9 | ||||||||||||||
| Financial Condition | |||||||||||||||||||
| Cash, cash equivalents and marketable securities | $ | 3,316.0 | $ | 1,848.5 | $ | 3,742.4 | $ | 3,107.4 | $ | 1,950.8 | |||||||||
| Total assets | $ | 14,316.6 | $ | 11,863.3 | $ | 10,130.1 | $ | 9,049.6 | $ | 8,092.5 | |||||||||
| Notes payable, line of credit and other financing arrangements, less current portion | $ | 582.1 | $ | 592.4 | $ | 687.4 | $ | 1,060.8 | $ | 1,066.4 | |||||||||
| Total Biogen Idec Inc. shareholders’ equity | $ | 10,809.0 | $ | 8,620.2 | $ | 6,961.5 | $ | 6,425.5 | $ | 5,396.5 |
In addition to the following notes, the financial data included within the tables above should be read in conjunction with our consolidated financial statements and related notes and the “Management’s Discussion and Analysis of Financial Condition and Results of Operations” sections of this report and our previously filed Form 10-Ks.
| (1) | Included in total cost and expenses are charges to acquired in-process research and development (IPR&D) totaling $245.0 million. Of this amount, $205.0 million was incurred in connection with the license agreement entered into with Knopp Neurosciences Inc. (Knopp), which we consolidated as we determined that we were the primary beneficiary of the entity. The $205.0 million charge was partially offset by an attribution of $145.0 million to the noncontrolling interest. We also incurred a charge of $40.0 million in connection with our acquisition of Biogen Idec Hemophilia Inc. (BIH), formerly Syntonix, related to the initiation of patient enrollment in a registrational trial of ALPROLIX. |
| (2) | Our share of revenues from unconsolidated joint business reflects charges of $50.0 million in 2011 and $49.7 million in 2013 for damages and interest awarded to Hoechst in Genentech's arbitration with Hoechst for RITUXAN. |
| (3) | Biogen Idec Inc.’s shareholders’ equity reflects a reduction in additional paid in capital and noncontrolling interests totaling $187.3 million resulting from our purchase of the noncontrolling interest in our joint venture investments in Biogen Dompé SRL and Biogen Dompé Switzerland GmbH. |
| (4) | Commencing in the second quarter of 2013, product and total revenues include 100% of net revenues related to sales of TYSABRI as a result of our acquisition of all remaining rights to TYSABRI from Elan and net revenues related to sales of TECFIDERA. In addition, upon the closing of our acquisition of all remaining rights to TYSABRI, our collaboration agreement was terminated, and we no longer record collaboration profit sharing. |
Item 7. Management’s Discussion and Analysis of Financial Condition and Results of Operations
The following discussion should be read in conjunction with our consolidated financial statements and related notes beginning on page F-1 of this report. Certain totals may not sum due to rounding.
Executive Summary
Introduction
Biogen Idec is a global biopharmaceutical company focused on discovering, developing, manufacturing and delivering therapies for neurological, autoimmune and hematologic disorders. Our principal marketed products include AVONEX, PLEGRIDY, TECFIDERA, TYSABRI, and FAMPYRA for multiple sclerosis (MS), ALPROLIX for hemophilia B and ELOCTATE for hemophilia A. We also collaborate on the development and commercialization of RITUXAN for the treatment of non-Hodgkin's lymphoma, chronic lymphocytic leukemia and other conditions and share profits and losses for GAZYVA which is approved for the treatment of chronic lymphocytic leukemia.
Our current revenues depend upon continued sales of our principal products. We may be substantially dependent on sales from our principal products for many years, including an increasing reliance on sales of TECFIDERA as we expand into additional markets. In the longer term, our revenue growth will be dependent upon the successful clinical development, regulatory approval and launch of new commercial products, our ability to obtain and maintain patents and other rights related to our marketed products and assets originating from our research and development efforts, and successful execution of external business development opportunities. As part of our ongoing research and development efforts, we have devoted significant resources to conducting clinical studies to advance the development of new pharmaceutical products and to explore the utility of our existing products in treating disorders beyond those currently approved in their labels.
Financial Highlights
The following table is a summary of financial results achieved:
| For the Years Ended December 31, | % Change | |||||||||
| 2014 compared to 2013 | ||||||||||
| (In millions, except per share amounts and percentages) | 2014 | 2013 (1) | ||||||||
| Total revenues | $ | 9,703.3 | $ | 6,932.2 | 40.0 | % | ||||
| Income from operations | $ | 3,972.4 | $ | 2,515.5 | 57.9 | % | ||||
| Net income attributable to Biogen Idec Inc. | $ | 2,934.8 | $ | 1,862.3 | 57.6 | % | ||||
| Diluted earnings per share attributable to Biogen Idec Inc. | $ | 12.37 | $ | 7.81 | 58.3 | % |
| (1) | Commencing in the second quarter of 2013, product and total revenues include 100% of net revenues related to TYSABRI as a result of our acquisition of all remaining rights to TYSABRI from Elan and net revenues related to sales of TECFIDERA. |
As described below under “Results of Operations,” our income from operations for the year ended December 31, 2014, reflect the following:
| • | Worldwide AVONEX revenues totaled $3,013.1 million for 2014, representing an increase of 0.3% over 2013. |
| • | Worldwide PLEGRIDY revenues totaled $44.5 million for 2014. |
| • | Worldwide TECFIDERA revenues totaled $2,909.2 million for 2014, representing an increase of 232.1% over 2013. |
| • | Worldwide TYSABRI revenues totaled $1,959.5 million for 2014, representing an increase of 28.4% over 2013. |
| • | Worldwide FAMPYRA revenues totaled $80.2 million for 2014, representing an increase of 8.4% over 2013. |
| • | Worldwide ALPROLIX revenues totaled $76.0 million for 2014. |
| • | Worldwide ELOCTATE revenues totaled $58.4 million for 2014. |
| • | Our share of revenues from unconsolidated joint business totaled $1,195.4 million for 2014, representing an increase of 6.2% from 2013. |
| • | Total cost and expenses increased 29.4% for 2014 compared to 2013. This increase resulted from a 42.8% increase in the amortization of acquired intangible assets, a 36.5% increase in cost of sales, a 31.1% increase in research and development expense and a 30.4% increase in selling, general and administrative expense, partially offset by a 100.0% decrease in collaboration profit sharing and an increase in the gain on fair value remeasurement of contingent consideration compared with the same period in 2013. |
The change in amortization of acquired intangible assets was primarily driven by a $60.2 million increase in amortization of acquired and in-licensed rights and patents as we recognized a full year of expense related to our TYSABRI rights in 2014 versus nine months of expense in 2013, total impairment charges of $50.9 million related to one of our out-licensed patents and one of our in-process research and development intangible assets, and higher amortization of our developed technology intangible asset as a result of lower expected lifetime revenues of AVONEX versus the prior year.
The increase in cost of sales was primarily driven by higher unit sales volume, including recent product launches and higher royalty payments due to Elan. The increase in research and development expense was primarily related to an increase in costs incurred in connection with our early stage programs and milestone and upfront payments. Higher selling, general and administrative expense resulted from increased costs incurred in connection with our recent product launches.
We generated $2,942.1 million of net cash flows from operations for 2014, which were primarily driven by earnings offset by an increase in working capital. Cash, cash equivalents and marketable securities totaled approximately $3,316.0 million as of December 31, 2014.
Business Environment
We conduct our business within the biopharmaceutical industry, which is highly competitive. Many of our competitors are working to develop or have commercialized products similar to those we market or are developing. In addition, the commercialization of certain of our own approved MS products and pipeline product candidates may negatively impact future sales of our MS products. Our products may also face increased competitive pressures from the introduction of generic versions, related prodrug derivatives or biosimilars of existing products and other technologies, such as gene therapies.
The Patient Protection and Affordable Care Act (PPACA)
The PPACA included a significant expansion of the Medicaid program, as well as the creation of new state-based health benefit exchanges, or marketplaces, through which individuals and small businesses may purchase health insurance. Premium and cost-sharing credits and subsidies are available to those who qualify based on income. Marketplace plans began to enroll new members in October 2013, and coverage began on January 1, 2014. Although the effects of the legislation are still unclear, PPACA will result in a greater number of individuals with health insurance under Medicaid and the marketplace health plans. The impact on manufacturers, including us, will depend in part on the formulary and benefit design decisions made by insurance sponsors or plans participating in the programs. It is possible that individuals who were previously unable to access insurance may now become insured, thus increasing coverage for our products. This potential increase in coverage, however, may be offset by the added discounts that could be required in these channels as well as the number of patients who over time move from commercial insurance to the health insurance marketplaces. It is also possible that we may need to provide discounts or rebates to such plans in order to maintain favorable formulary access for our products for this patient population, which could have an adverse impact on our sales
Showing the first 8K of 132K characters. Open the full section
Item 7A. Quantitative and Qualitative Disclosures About Market Risk
The information required by this Item is incorporated by reference to the discussion under “Market Risk” in Item 7. “Management’s Discussion and Analysis of Financial Condition and Results of Operations.”
Item 8. Financial Statements and Supplementary Data
The information required by this Item 8 is contained on pages F-1 through F-65 of this report and is incorporated herein by reference.
Item 9. Changes in and Disagreements with Accountants on Accounting and Financial Disclosure
None.
Item 9A. Controls and Procedures
Disclosure Controls and Procedures and Internal Control over Financial Reporting
Controls and Procedures
We have carried out an evaluation, under the supervision and with the participation of our management, including our principal executive officer and principal financial officer, of the effectiveness of the design and operation of our disclosure controls and procedures (as defined in Rules 13a-15(e) and 15d-15(e) under the Securities Exchange Act of 1934, as amended), as of December 31, 2014. Based upon that evaluation, our principal executive officer and principal financial officer concluded that, as of the end of the period covered by this report, our disclosure controls and procedures are effective in ensuring that (a) the information required to be disclosed by us in the reports that we file or submit under the Securities Exchange Act is recorded, processed, summarized and reported within the time periods specified in the SEC’s rules and forms, and (b) such information is accumulated and communicated to our management, including our principal executive officer and principal financial officer, as appropriate to allow timely decisions regarding required disclosure. In designing and evaluating our disclosure controls and procedures, our management recognized that any controls and procedures, no matter how well designed and operated, can provide only reasonable assurance of achieving the desired control objectives, and our management necessarily was required to apply its judgment in evaluating the cost-benefit relationship of possible controls and procedures.
Changes in Internal Control over Financial Reporting
There were no changes in our internal control over financial reporting during the quarter ended December 31, 2014 that have materially affected, or are reasonably likely to materially affect, our internal control over financial reporting.
Management’s Annual Report on Internal Control over Financial Reporting
Our management is responsible for establishing and maintaining adequate internal control over our financial reporting. Internal control over financial reporting is defined in Rules 13a-15(f) and 15d-15(f) under the Securities Exchange Act as a process designed by, or under the supervision of, a company’s principal executive and principal financial officers and effected by a company’s board of directors, management and other personnel to provide reasonable assurance regarding the reliability of financial reporting and the preparation of financial statements for external purposes in accordance with U.S. GAAP. Our internal control over financial reporting includes those policies and procedures that:
| • | pertain to the maintenance of records that, in reasonable detail, accurately and fairly reflect our transactions and dispositions of our assets; |
| • | provide reasonable assurance that transactions are recorded as necessary to permit preparation of financial statements in accordance with U.S. GAAP, and that our receipts and expenditures are being made only in accordance with authorizations of our management and directors; and |
| • | provide reasonable assurance regarding prevention or timely detection of unauthorized acquisition, use or disposition of our assets that could have a material effect on our financial statements. |
Because of its inherent limitations, internal control over financial reporting may not prevent or detect misstatements. Projections of any evaluation of effectiveness to future periods are subject to the risk that controls may become inadequate because of changes in conditions, or that the degree of compliance with the policies or procedures may deteriorate.
Our management assessed the effectiveness of our internal control over financial reporting as of December 31, 2014. In making this assessment, management used the criteria set forth by the Committee of Sponsoring Organizations of the Treadway Commission (COSO) in its 2013 Internal Control — Integrated Framework.
Based on our assessment, our management has concluded that, as of December 31, 2014, our internal control over financial reporting is effective based on those criteria.
The effectiveness of our internal control over financial reporting as of December 31, 2014 has been audited by PricewaterhouseCoopers LLP, an independent registered public accounting firm, as stated in their attestation report, which is included herein.
Item 9B. Other Information
None.
PART III
Item 10. Directors, Executive Officers and Corporate Governance
The information concerning our executive officers is set forth under the heading “Our Executive Officers” in Part I of this report. The text of our code of business conduct, which includes the code of ethics that applies to our principal executive officer, principal financial officer, principal accounting officer or controller, and persons performing similar functions, is posted on our website, www.biogenidec.com, under the “Corporate Governance” subsection of the “About Us” section of the site. We intend to make all required disclosures regarding any amendments to, or waivers from, provisions of our code of business conduct at the same location of our website.
The response to the remainder of this item is incorporated by reference from the discussion responsive thereto in the sections entitled “Proposal 1 - Election of Directors,” “Corporate Governance,” “Stock Ownership - Section 16(a) Beneficial Ownership Reporting Compliance” and “Miscellaneous - Stockholder Proposals” contained in the proxy statement for our 2015 annual meeting of stockholders.
Item 11. Executive Compensation
The response to this item is incorporated by reference from the discussion responsive thereto in the sections entitled “Executive Compensation and Related Information” and “Corporate Governance” contained in the proxy statement for our 2015 annual meeting of stockholders.
Item 12. Security Ownership of Certain Beneficial Owners and Management and Related Stockholder Matters
The response to this item is incorporated by reference from the discussion responsive thereto in the sections entitled “Stock Ownership” and “Equity Compensation Plan Information” contained in the proxy statement for our 2015 annual meeting of stockholders.
Item 13. Certain Relationships and Related Transactions, and Director Independence
The response to this item is incorporated by reference from the discussion responsive thereto in the sections entitled “Certain Relationships and Related Person Transactions” and “Corporate Governance” contained in the proxy statement for our 2015 annual meeting of stockholders.
Item 14. Principal Accountant Fees and Services
The response to this item is incorporated by reference from the discussion responsive thereto in the section entitled “Proposal 2 — Ratification of the Selection of our Independent Registered Public Accounting Firm” contained in the proxy statement for our 2015 annual meeting of stockholders.
PART IV
Item 15. Exhibits and Financial Statement Schedules
| a. | (1) Consolidated Financial Statements: |
The following financial statements are filed as part of this report:
| Financial Statements | Page Number | |
| Consolidated Statements of Income | F-2 | |
| Consolidated Statements of Comprehensive Income | F-3 | |
| Consolidated Balance Sheets | F-4 | |
| Consolidated Statements of Cash Flows | F-5 | |
| Consolidated Statements of Equity | F-6 | |
| Notes to Consolidated Financial Statements | F-9 | |
| Report of Independent Registered Public Accounting Firm | F-64 |
(2) Financial Statement Schedules
Schedules are omitted because they are not applicable, or are not required, or because the information is included in the consolidated financial statements and notes thereto.
(3) Exhibits
The exhibits listed on the Exhibit Index beginning on page A-1, which is incorporated herein by reference, are filed or furnished as part of this report or are incorporated into this report by reference.
SIGNATURES
Pursuant to the requirements of Section 13 or 15(d) of the Securities Exchange Act of 1934, the registrant has duly caused this report to be signed on its behalf by the undersigned, thereunto duly authorized.
| BIOGEN IDEC INC. | |
| By: | /S/ GEORGE A. SCANGOS |
| George A. Scangos | |
| Chief Executive Officer |
Date: February 4, 2015
Pursuant to the requirements the Securities Exchange Act of 1934, this report has been signed below by the following persons on behalf of the registrant and in the capacities and on the dates indicated.
| Name | Capacity | Date | ||
| /S/ GEORGE A. SCANGOS | Director and Chief Executive Officer (principal executive officer) | February 4, 2015 | ||
| George A. Scangos | ||||
| /S/ PAUL J. CLANCY | Executive Vice President, Finance and Chief Financial Officer (principal financial officer) | February 4, 2015 | ||
| Paul J. Clancy | ||||
| /S/ GREGORY F. COVINO | Vice President, Finance, Chief Accounting Officer (principal accounting officer) | February 4, 2015 | ||
| Gregory F. Covino | ||||
| /S/ STELIOS PAPADOPOULOS | Director and Chairman of the Board of Directors | February 4, 2015 | ||
| Stelios Papadopoulos | ||||
| /S/ ALEXANDER J. DENNER | Director | February 4, 2015 | ||
| Alexander J. Denner | ||||
| /S/ CAROLINE D. DORSA | Director | February 4, 2015 | ||
| Caroline D. Dorsa | ||||
| /S/ NANCY L. LEAMING | Director | February 4, 2015 | ||
| Nancy L. Leaming | ||||
| /S/ RICHARD C. MULLIGAN | Director | February 4, 2015 | ||
| Richard C. Mulligan | ||||
| /S/ ROBERT W. PANGIA | Director | February 4, 2015 | ||
| Robert W. Pangia | ||||
| /S/ BRIAN S. POSNER | Director | February 4, 2015 | ||
| Brian S. Posner | ||||
| /S/ ERIC K. ROWINSKY | Director | February 4, 2015 | ||
| Eric K. Rowinsky | ||||
| /S/ LYNN SCHENK | Director | February 4, 2015 | ||
| Lynn Schenk | ||||
| /S/ STEPHEN A. SHERWIN | Director | February 4, 2015 | ||
| Stephen A. Sherwin |
BIOGEN IDEC INC. AND SUBSIDIARIES
CONSOLIDATED FINANCIAL STATEMENTS
| Page Number | ||
| Consolidated Statements of Income | F-2 | |
| Consolidated Statements of Comprehensive Income | F-3 | |
| Consolidated Balance Sheets | F-4 | |
| Consolidated Statements of Cash Flows | F-5 | |
| Consolidated Statements of Equity | F-6 | |
| Notes to Consolidated Financial Statements | F-9 | |
| Report of Independent Registered Public Accounting Firm | F-64 |
F- 1
BIOGEN IDEC INC. AND SUBSIDIARIES
CONSOLIDATED STATEMENTS OF INCOME
(In thousands, except per share amounts)
| For the Years Ended December 31, | |||||||||||
| 2014 | 2013 | 2012 | |||||||||
| Revenues: | |||||||||||
| Product, net | $ | 8,203,404 | $ | 5,542,331 | $ | 4,166,074 | |||||
| Unconsolidated joint business | 1,195,389 | 1,126,017 | 1,137,923 | ||||||||
| Other | 304,531 | 263,851 | 212,464 | ||||||||
| Total revenues | 9,703,324 | 6,932,199 | 5,516,461 | ||||||||
| Cost and expenses: | |||||||||||
| Cost of sales, excluding amortization of acquired intangible assets | 1,171,036 | 857,726 | 545,494 | ||||||||
| Research and development | 1,893,422 | 1,444,053 | 1,334,919 | ||||||||
| Selling, general and administrative | 2,232,342 | 1,712,051 | 1,277,465 | ||||||||
| Amortization of acquired intangible assets | 489,761 | 342,948 | 202,204 | ||||||||
| Collaboration profit sharing | — | 85,357 | 317,895 | ||||||||
| (Gain) loss on fair value remeasurement of contingent consideration | (38,893 | ) | (547 | ) | 27,202 | ||||||
| Restructuring charges | — | — | 2,225 | ||||||||
| Total cost and expenses | 5,747,668 | 4,441,588 | 3,707,404 | ||||||||
| Gain on sale of rights | 16,758 | 24,898 | 46,792 | ||||||||
| Income from operations | 3,972,414 | 2,515,509 | 1,855,849 | ||||||||
| Other income (expense), net | (25,781 | ) | (34,930 | ) | (744 | ) | |||||
| Income before income tax expense and equity in loss of investee, net of tax | 3,946,633 | 2,480,579 | 1,855,105 | ||||||||
| Income tax expense | 989,942 | 601,014 | 470,554 | ||||||||
| Equity in loss of investee, net of tax | 15,126 | 17,224 | 4,518 | ||||||||
| Net income | 2,941,565 | 1,862,341 | 1,380,033 | ||||||||
| Net income attributable to noncontrolling interests, net of tax | 6,781 | — | — | ||||||||
| Net income attributable to Biogen Idec Inc. | $ | 2,934,784 | $ | 1,862,341 | $ | 1,380,033 | |||||
| Net income per share: | |||||||||||
| Basic earnings per share attributable to Biogen Idec Inc. | $ | 12.42 | $ | 7.86 | $ | 5.80 | |||||
| Diluted earnings per share attributable to Biogen Idec Inc. | $ | 12.37 | $ | 7.81 | $ | 5.76 | |||||
| Weighted-average shares used in calculating: | |||||||||||
| Basic earnings per share attributable to Biogen Idec Inc. | 236,359 | 236,919 | 237,938 | ||||||||
| Diluted earnings per share attributable to Biogen Idec Inc. | 237,176 | 238,308 | 239,740 |
See accompanying notes to these consolidated financial statements.
F- 2
BIOGEN IDEC INC. AND SUBSIDIARIES
CONSOLIDATED STATEMENTS OF COMPREHENSIVE INCOME
(In thousands)
| For the Years Ended December 31, | |||||||||||
| 2014 | 2013 | 2012 | |||||||||
| Net income attributable to Biogen Idec Inc. | $ | 2,934,784 | $ | 1,862,341 | $ | 1,380,033 | |||||
| Other comprehensive income: | |||||||||||
| Unrealized gains (losses) on securities available for sale: | |||||||||||
| Unrealized gains (losses) recognized during the period, net of tax of $80, $6,394 and $2,940 | 411 | 11,770 | 5,080 | ||||||||
| Less: reclassification adjustment for (gains) losses included in net income, net of tax of $3,462, $5,576 and $486 | (6,429 | ) | (10,355 | ) |
Showing the first 8K of 285K characters. Open the full section